

Submitted:
07 October 2026
Posted:
09 October 2026
You are already at the latest version
Abstract
Population genomics can improve mosquito control when genetic measurements are linked to specific design, deployment, and monitoring decisions. Here, we provide a decision-oriented framework for integrating population genetic data into conventional sterile insect technique (SIT), female-specific Release of Insects carrying a Dominant Lethal (fsRIDL), CRISPR-based precision-guided SIT (pgSIT), gene drives, Wolbachia-based incompatible insect technique (IIT), and Wolbachia population modification. These approaches have distinct biological requirements and failure modes. Immigration can replenish vector populations targeted by repeated sterile male or IIT releases; standing or newly generated functional escape alleles can impede the performance of homing gene drives; and Wolbachia establishment depends on cytoplasmic incompatibility, maternal transmission, host fitness, and spatial structure. We show how target site surveys, functional and compatibility assays, movement measurements, contained experiments, and spatial models can inform target selection, release stock preparation, release geography, and surveillance. We distinguish evidence from field trials—including mosquito suppression with IIT and reductions in dengue following IIT-SIT or Wolbachia population-modification releases—from gene-drive outcomes that remain pre-field. We argue that population genetic measurements are most useful when they lead to prespecified, testable decisions: local inputs should define forecast ranges, endpoints, and decision rules that can be evaluated against independent observations. Genomic structure alone cannot establish current movement, biological compatibility, field efficacy, or epidemiological benefit. Linking population genetic measurements to intervention-specific decisions and prospective validation provides a practical framework for translating mosquito genetic control from laboratory design toward responsible field deployment.
Keywords:
population genomics
; mosquito control
; Wolbachia
; incompatible insect technique
; population modification
; gene drive
; resistance evolution
; spatial population structure
; genomic surveillance
1. Introduction
Mosquito-borne pathogens, including those responsible for malaria, dengue, Zika, chikungunya, and yellow fever, remain a major global public-health burden, particularly where vector control is central to disease prevention [1,2,3,4]. Insecticide-treated nets, indoor residual spraying, ultra-low-volume adulticide and larvicide applications [5], and source reduction have reduced transmission in many settings, but their effectiveness can be constrained by insecticide resistance, operational challenges, urbanization, and environmental change [6,7]. Genetic and symbiont-based biocontrol approaches offer complementary strategies to suppress vector populations or reduce their capacity to transmit pathogens. Integrated vector management (IVM) provides a framework for selecting and coordinating control methods according to local vector ecology, epidemiological evidence, and operational resources [8]. Genetic and symbiont-based biocontrol approaches offer additional tools within this framework to suppress vector populations or reduce their capacity to transmit pathogens.
These include conventional sterile insect technique (SIT) [9,10], Female-Specific Release of Insects carrying a Dominant Lethal (fsRIDL) [11,12], CRISPR/Cas9-based precision-guided SIT (pgSIT) [13,14,15,16], gene drives [17,18], Wolbachia-based incompatible insect technique (IIT) for population suppression [19,20,21], and Wolbachia population modification for pathogen blocking [22]. Although molecular engineering and release-stock preparation are critical to these approaches, their performance in the field also depends on the genetic, ecological, and demographic characteristics of the populations in which they are deployed.
Wild mosquito populations vary in target sequences [23,24], resistance phenotypes, population structure [25,26] , and connectivity. Such variation can alter intervention outcomes even when a system performs well in the laboratory. Target-site polymorphisms may reduce CRISPR/Cas9 cleavage [27,28], population connectivity can influence spread, confinement, or immigration [29,30], and DNA repair coupled with selection can favor functional escape genotypes [28,31]. Laboratory colonies may capture only a fraction of this diversity [32], underscoring the need to evaluate candidate interventions in the context of genetically and geographically representative populations.
Previous reviews have examined genetic biocontrol technologies and resistance evolution [31,33,34], while Schmidt et al. surveyed broader applications of population genomics to mosquito invasions, insecticide resistance, and rear-and-release programs [35]. Here, we take a complementary, decision-oriented approach: rather than asking only what population genetic measurements can reveal, we ask how those measurements should change the design, deployment, and evaluation of specific mosquito control interventions. Target haplotypes can inform molecular design, connectivity can guide release geography and surveillance, and field measurements can test assumptions that genomic data alone cannot resolve. For Wolbachia-based approaches, infection dynamics, compatibility, and release stock background provide additional intervention-specific considerations [29,35,36]. This review therefore organizes evidence from population genomic surveys, functional assays, contained experiments, field releases, and models according to the decisions each can inform and the limitations of each evidence type. We distinguish modeled predictions from observed field outcomes and emphasize prospective validation in which local measurements define testable forecasts, endpoints, and decision rules. Figure 1 illustrates this measurement-to-decision framework for genetic control approaches, and Table 1 summarizes the architecture-specific measurements, evidence sources, evidence limitations, and illustrative decision triggers for the interventions examined in depth.
2. Standing Genetic Variation: Hidden Constraints on Targeting
Standing genetic variation can constrain target sequence selection used for a control before release. Wild mosquito populations harbor geographically heterogeneous target-site diversity [37,38,39,40], and even a single nucleotide change within a guide RNA binding site or adjacent motif can reduce CRISPR/Cas9 cleavage [40,41,42]. Resistant haplotypes may therefore predate an intervention and go undetected when constructs are designed and optimized in genetically narrow laboratory colonies [32,43,44]. Sequence variation alone, however, does not establish resistance. Whether a variant constitutes functional resistance depends on its position, guide-specific effects on cleavage, consequences for target-gene function, and associated fitness costs. In homing drives, alleles that reduce cleavage while preserving target-gene function can be favored by repeated cleavage and selection [28,45,46], whereas variants that disrupt an essential gene may carry substantial fitness costs. Reduced cleavage can impair drive conversion or homology-directed repair in homing systems. By contrast, pgSIT releases sterile males whose intended phenotype is generated before release and does not depend on continued cleavage or inheritance in the wild population; target site integrity in the wild population is therefore not a determinant of field efficacy. Conventional irradiation-based SIT likewise has no CRISPR target in the released population; its principal constraints instead include male competitiveness, release ratio, and immigration [47,48].
Design should therefore begin with geographically and seasonally-representative surveys of candidate target sites coupled with functional testing. Where feasible, interventions should favor conserved, functionally-constrained loci and incorporate multiple guides so that performance does not depend on a single cleavage site [14,15,16,37,40,49,50,51,52]. Comparative genomics and functional annotation can identify candidate loci that are conserved across wild populations, but sequence conservation is only the first step: field-derived haplotypes should subsequently be evaluated for cleavage, repair outcomes, gene function, phenotype, and fitness in relevant genetic backgrounds [14,15,16,37,40,41,49,50]. Laboratory and semi-field experiments can further reveal mating or performance costs that sequence data alone cannot detect [43,44,53].
Sampling depth and statistical power are equally important when interpreting apparent target conservation. Under ideal independent sampling, the probability of detecting at least one copy of a variant present at frequency p among n sampled alleles is . Thus, a variant present at 1% frequency has approximately a 63% probability of being observed among 100 sampled alleles and a 95% probability among 300. In practice, relatedness among samples, uneven geographic coverage, seasonality, sequencing depth, and assay error can further reduce detection probability. Surveys should therefore report the numbers of mosquitoes, alleles, and sites sampled; sampling dates and geographic coverage; quality-control criteria; and the variant frequencies they were adequately powered to detect. Failure to observe a variant in a single colony, locality, or season should not be interpreted as evidence of invariance. These measurements can guide target selection and define the initial genetic conditions for an intervention, while population movement determines whether locally observed variation remains localized or becomes a regional constraint (Box 1).
Box 1. Population-genetic measurements and their limits.
Genomic surveys can estimate target-site haplotype frequencies and the power to detect rare variants, while the fixation index (FST) describes genetic differentiation among sampled populations. Estimates of effective population size can help characterize the potential influence of genetic drift, whereas ancestry and admixture analyses can identify candidate population mixing. However, each measure depends on sampling scale, season, marker choice, and model assumptions. Effective population size can also be influenced by selection and migration, particularly in recently-invaded or insecticide-exposed populations, and should not be interpreted as census population size or as a direct measure of contemporary movement [25,35,54].
The appropriate genomic method should therefore follow the biological question. Genome-wide markers can resolve ancestry, population structure, and close-kin relationships; guide selection requires high-quality sequence information spanning the target site and adjacent motif; and copy-number variants or inversions require methods and reference assemblies capable of resolving structural variation [35]. Reduced-representation datasets generated using different protocols may share few markers [55], limiting comparisons across sites or studies [35]. Importantly, failure to detect a variant does not establish its absence from other locations, seasons, or unsampled populations. Likewise, FST describes genetic differentiation rather than contemporary movement, and evidence of admixture alone does not demonstrate fertile hybridization or viable introgression of an engineered construct.
Release-scale connectivity therefore requires complementary evidence from direct measurements of movement, mating, and temporal population dynamics [35,56,57,58,59,60,61]. Similarly, candidate low-cleavage variants should not be classified as functionally resistant based on sequence alone; cleavage, target-gene function, and fitness must be evaluated experimentally. Population-genetic summaries are therefore best used to identify what and where to sample and test, rather than as stand-alone evidence for defining release boundaries.
3. Gene Flow and Spatial Population Structure
Standing genetic variation establishes local initial conditions, whereas movement determines how broadly that variation is shared among populations. Anopheles and Aedes populations can exhibit fine-scale genetic differentiation over relatively short distances, allowing target-site, resistance, and modifier-allele frequencies to differ even among neighboring populations [56,57,58,59]. In the An. gambiae complex, there are a number of studies reported hybridization between An. gambiae s.s. and An. coluzzii which usually disappears likely due to relatively low fitness compared to parental species in nature [62]. However, adaptive introgression of insecticide resistance mutation transferred from An. gambiae s.s. to An. coluzzii followed by selection on standing variation of metabolic resistance gene altered local species composition and target haplotypes in West Africa [63,64,65]. The timing of the adaptive introgression and rise of insecticide resistant An. coluzzii coincides with scaled up distribution of insecticide treated bednets suggesting that human-assisted introduction of adaptive insecticide-resistance alleles into An. coluzzii [63]. In urban Ae. aegypti, fine-scale neighborhood structure can coexist with movement or human-assisted transport between otherwise differentiated populations [57,58,59,61]. Release boundaries inferred solely from local population structure may therefore overlook transport-linked introductions [66,67] , and sampling and surveillance should include plausible source populations and transport corridors where such movement is relevant. Importantly, none of these patterns provides a universal estimate of dispersal. Surveys should span relevant habitat nodes and seasons, distinguish contemporary adult movement from longer-term introgression, and characterize connectivity among populations relevant to release and surveillance planning. Figure 2 illustrates how species composition, local movement, and transport-linked connectivity can define different sampling and release questions in Anopheles and Aedes.
Spatial structure can influence both intervention performance and deployment strategy. Target polymorphisms, genetic background, or expression context in field populations may differ from those in laboratory strains, while dispersal can spread a self-sustaining drive, introduce resistant alleles, or replenish populations undergoing self-limiting suppression [10,28,47,48,68,69,70,71]. Close-kin mark-recapture methods recovered dispersal and barrier parameters in simulated Ae. aegypti populations [72], while behavioral-state simulations demonstrate how resource-seeking behavior can generate spatially heterogeneous movement [73]. These results illustrate approaches for characterizing movement rather than providing transferable field dispersal estimates. Consistent with this need for local measurement, a field study of Ae. aegypti in a Malaysian apartment complex found that close-kin pairs were concentrated within the same or adjacent buildings, demonstrating fine-scale spatial structure relevant to release planning [35,74].
The consequences of connectivity also depend on intervention architecture. In continuous habitat, homing drives may spread as spatial waves, whereas fragmented habitat, source–sink dynamics, and asymmetric movement can produce heterogeneous establishment or local fade-out [68,75,76,77]. Threshold-dependent systems may fail to establish when local introduction frequencies remain below the required threshold [78,79,80]. Split-drive components occupy separate loci, limiting sustained propagation, although predicted confinement remains dependent on local demography and movement [79,81,82,83]. Highly connected populations may therefore require broader release and surveillance coverage, whereas more isolated populations may permit more localized strategies. Surveillance for resistance should likewise include populations connected to release sites by relevant migration pathways. For IIT, immigration can replenish fertile females following suppression [19,20,21]; for Wolbachia population modification, dispersal and local infection frequencies influence establishment and subsequent spatial spread [22,29,35]. Genomic data can identify candidate connections among populations, but release-scale connectivity, reproductive compatibility, and maternal transmission require direct measurement such as mark release and re-capture.
4. Resistance Evolution
Resistance is an expected evolutionary response to strong selection, but its origin, trajectory, and consequences depend on both the intervention architecture and the population in which it operates. In CRISPR/Cas9 gene drive systems, escape alleles can arise from standing target-site variation, de novo mutation, or repair following cleavage [28,37,40,41,84]. Error-prone nonhomologous end joining (NHEJ) can generate insertions or deletions that disrupt guide recognition or the adjacent motif, whereas homology-directed repair (HDR) can copy the drive; the relative frequency of these outcomes depends on nuclease expression and repair context [28,85,86,87,88,89]. Large populations also provide greater mutational opportunity, allowing initially rare functional escape alleles to become relevant under sustained selection [28]. Variants that reduce or prevent cleavage while preserving target gene function and fitness are particularly important because they can increase under drive-mediated selection. Their trajectories depend on selection strength, dominance, and fitness costs: dominant or partially dominant alleles are exposed to selection in heterozygotes, whereas recessive alleles may remain phenotypically masked until homozygous. Genetic drift becomes increasingly important when alleles are rare or suppression reduces population size, while migration can introduce or reseed resistance across connected populations.
Contained studies demonstrate that resistance outcomes are system-specific rather than governed by a universal time to failure. Resistance reduced drive performance in some cage experiments and models [88,89,90,91,92,93], whereas a doublesex-targeting suppression drive eliminated age-structured large cage populations without selecting functional resistance [94]. In contrast, a resistant allele accumulated in one of several two-year Anopheles gambiae population-modification cage replicates [95]. Morianou and colleagues provide a particularly clear measurement-to-redesign example [96], combining surveys of standing and drive-induced doublesex variants with functional testing, modeling, target redesign, and laboratory validation of multiplexed drives. Together, these studies illustrate why sequence variation must be linked to functional evidence before an allele is classified as resistant and why predicted robustness at natural-population scales requires prospective validation.
Resistance mechanisms also differ fundamentally among intervention architectures. Irradiation-based SIT has no CRISPR target and no sustained inheritance, although male competitiveness, immigration, and the need for repeated releases remain important determinants of suppression [10,47,48]. Similarly, pgSIT releases sterile males whose phenotype is generated before release and does not depend on continued cleavage or inheritance in the wild population. For homing gene drives, resistance alleles can be classified as R1, R2, or R3: R1 alleles are cleavage-resistant variants that preserve target-gene function, R2 alleles are cleavage-resistant variants that disrupt target-gene function, and R3 alleles remain susceptible to cleavage but are not efficiently converted to the drive allele by homology-directed repair. These R1–R3 resistance classes are specific to homing gene drives and do not apply to SIT, fsRIDL, pgSIT, or Wolbachia-based approaches. Confinable drive architectures require system-specific assessment of target variation, repair outcomes, and persistence [97,98]. Across propagating or partially propagating systems, selection intensity, release strategy, migration, mutation supply, and demography jointly determine whether resistance emerges and persists.
For homing drives, several design strategies can reduce the number of viable evolutionary escape routes. Multiplexing guides and targeting conserved, functionally-constrained sequences can reduce the probability that a single functional variant prevents cleavage [51,52,99]. Cleave-and-rescue (ClvR) and homing-based rescue architectures, including the home-and-rescue (HomeR) system [100], can couple organismal viability to the engineered construct by targeting an essential gene while providing a recoded rescue, thereby reducing the selective advantage of nonfunctional resistant alleles. Germline-restricted or appropriately-timed nuclease activity aims to favor HDR over NHEJ, while haplolethal targets and reduced maternal carryover provide additional strategies [90,91,101,102,103,104,105]. The effectiveness of each approach must nevertheless be established experimentally for the specific construct and genetic background. Monitoring should also distinguish functional resistance from immigration, fitness costs, or other processes that can reduce construct frequency; no drive architecture should be assumed to be resistance-proof.
Insecticide resistance provides a useful parallel for genomic surveillance, rather than a mechanistic model of CRISPR drive escape. As reviewed by Schmidt et al. [35], insecticide resistance can involve interacting target site alleles, metabolic gene copy number changes, structural variants, or adaptive alleles introduced from other populations. A single diagnostic marker may therefore fail to capture the relevant phenotype or distinguish locally evolved resistance from immigration. For example, a recent genomic surveillance of voltage-gated sodium channel gene in Ae. aegypti in Florida populations revealed two new mutations as well as four mutations reported for the first time in Florida [106]. Some of these mutations were present in high frequencies (>18% resistant allele frequency) depending on specific county, suggesting that these mutations had been introduced or arisen a while ago and remain undetected due to limitation of surveillance method previously used.
Pre- and postintervention genomic sampling should be paired with insecticide bioassays and, where appropriate, functional assays, while ancestry and linked haplotypes can help distinguish introduction from local emergence. These measurements can inform insecticide-management and co-deployment decisions. By contrast, resistance to homing drives requires architecture-specific evidence linking target-site variation to cleavage, inheritance, target gene function, and fitness.
5. Wolbachia-Based Control: IIT and Population Modification
5.1. Incompatible Insect Technique (IIT)
Wolbachia-based incompatible insect technique (IIT) is a self-limiting population suppression strategy in which males carrying a Wolbachia strain that is incompatible with the local female population are repeatedly released. Matings between released males and incompatible females produce nonviable embryos through cytoplasmic incompatibility, and the released infection is not intended to spread through the target population. Suppression therefore depends on sustained release of competitive males and can be offset by immigration of fertile females [107,108]. A distinct operational risk is the accidental release of fertile Wolbachia-infected females, which can establish the released strain in the field and generate compatible offspring, potentially compromising the effectiveness of subsequent IIT releases. Strain-specific crossing tests, male mating competitiveness, sex-sorting accuracy [109], and surveillance for inadvertently released infected females are therefore central design and quality-control requirements. In combined IIT-SIT approaches, low-dose irradiation of residual females can reduce the risk of Wolbachia establishment, although the effects of irradiation and mass rearing on male quality and competitiveness still require measurement [20,21]. Field studies demonstrate suppression across several deployment settings. Standalone IIT suppressed Ae. albopictus at a subtropical field site [19], while combined IIT-SIT suppressed Ae. albopictus in Guangzhou [21] and reduced Ae. aegypti abundance and dengue risk in a cluster-randomized trial in Singapore [20].
5.2. Wolbachia Population Modification
Wolbachia population replacement modification involves releasing infected females and males so that a maternally-transmitted, pathogen-blocking infection can increase in frequency and persist in the target population. Cytoplasmic incompatibility can provide infected females with a reproductive advantage, but establishment and subsequent spatial spread depend on local infection frequency, maternal transmission fidelity, mosquito fitness, insecticide exposure, temperature, and movement [29,35,36]. Genomic surveys can characterize host ancestry, population structure, dispersal barriers, and the genetic similarity between the release stain and local populations, but the key intervention phenotypes require direct measurement. These include infection prevalence, cytoplasmic incompatibility, maternal transmission, pathogen blocking, and mosquito fitness. In a cluster-randomized trial in Yogyakarta. releases of wMel-infected Ae. aegypti reduced symptomatic, virologically confirmed dengue, with an estimated protective efficacy of 77.1% [22]. This result demonstrates epidemiological efficacy in the tested setting but should not be assumed to predict equivalent effects for other Wolbachia strains, mosquito populations, or deployment settings. Temperature can affect Wolbachia density and maternal transmission in Ae. aegypti, with responses differing among strains [110]. Field observations in Cairns also documented temporary declines in wMel infection frequency and density following a heatwave, followed by recovery [111].
These differences in mechanism define distinct checkpoints for Wolbachia-based interventions. IIT requires evidence of incompatible matings, competitive released males, and reliable exclusion of fertile infected females during repeated releases; population modification instead requires evidence that the infection establishes and persists, maintains effective maternal transmission, and reduces pathogen transmission. Host population genomics can guide sampling, identify potential barriers to spread, and inform release-strain selection, but these operational and epidemiological endpoints cannot be inferred from genomic data alone. Table 1 links each intervention architecture to the measurements and decision triggers that should be evaluated locally.
6. Integrating Population Genomics into System Design
Population genomics is most useful for system design when a measured pattern can be linked to a specific design or deployment decision. A locally common low-cleavage haplotype, for example, can motivate target redesign, whereas evidence of population connectivity can alter release geography or surveillance coverage [83]. Repair-outcome and machine learning models can help prioritize variants for experimental testing [112,113,114,115,116,117], but predictions generated in another species or genetic background should not be interpreted as field resistance probabilities. The relevant measurements also depend on intervention architecture. For IIT, compatibility and sex-sorting performance require direct testing [19,20,21]. For Wolbachia population modification, host ancestry can inform strain matching and spatial sampling, whereas infection frequency, maternal transmission, and pathogen blocking require direct measurement [22,29,36]. For homing drives, candidate resistance variants likewise require functional testing of cleavage, inheritance, target-gene function, and fitness before they can be classified as escape alleles. Figure 3 illustrates this measurement-to-outcome framework, distinguishing R1 alleles that block cleavage while retaining target gene function, R2 alleles that block cleavage but impair target-gene function, and R3 alleles that remain cleavable but are inefficiently converted to the drive allele. The qualitative scenarios further illustrate how these resistance classes could produce different trajectories for population suppression or transmission reduction in population-replacement strategies, while effector resistance could restore pathogen transmission even if the replacement drive itself persists. These resistance classes and their consequences are specific to homing-drive systems and should not be generalized to self-limiting or Wolbachia-based approaches. Each design or deployment decision should therefore be linked to a locally defined measurement and decision trigger, with measurement uncertainty propagated into setting-specific simulations [118].
The upper panel summarizes the classification framework using the Ag(QFS)1 Anopheles gambiae example [96]. Candidate field- or repair-derived variants are evaluated for effects on drive cleavage and biased inheritance, followed by assays of target-gene function, fertility, viability, and fitness. R1 alleles are cleavage-resistant variants that retain target-gene function and may therefore increase under drive-mediated selection and displace the drive; R2 alleles are cleavage-resistant variants that impair target-gene function and may be constrained by associated fitness costs; and R3 alleles remain susceptible to cleavage but are inefficiently converted to the drive allele by homology-directed repair, allowing drive inheritance while potentially reducing control efficacy. The lower panels illustrate qualitative consequences of these resistance classes for suppression and population-replacement drives. All quantities are normalized to their pre-intervention values and the trajectories are schematic rather than fitted predictions. For suppression drives, colored curves represent both relative total population size and proportional transmission potential, whereas the gray line represents the no-suppression reference. For replacement drives, total population size remains constant (gray), and colored curves represent transmission potential under the indicated resistance scenarios. The dotted magenta curve illustrates pathogen escape from the effector despite persistence of the replacement drive. Other determinants of transmission are held constant. These scenarios are illustrative proxies, not experimental results or predictions for particular releases. R1–R3 describe resistance to homing gene drives and do not apply to SIT, fsRIDL, pgSIT, or Wolbachia-based approaches.
An operational pipeline for a candidate An. gambiae homing-drive guide, for example, could begin by estimating target site and surrounding haplotype frequencies from geographically representative genomes [37,40]. Common local variants could then be tested for cleavage, repair outcomes, target gene function, and fitness. A spatial model could integrate these measurements with dispersal, seasonality, release size, and density dependence to evaluate establishment and resistance trajectories across plausible parameter ranges [68,77]. A common low-cleavage, function-preserving haplotype could trigger selection of a different guide or addition of another target, whereas uncertainty dominated by dispersal would instead prioritize movement measurements and refinement of release and surveillance boundaries. Independent cage populations could then test prespecified predictions before field evaluation. This workflow is an illustrative synthesis of separate evidence streams rather than a validated forecast for any particular site.
For self-limiting suppression, the decision framework is different. In Ae. aegypti, repeated sampling across neighborhoods and transport corridors [57,58,59,61] could identify population connections through which immigration may replenish a suppressed population. Direct measurements of movement, wild population abundance, and mating performance could then inform release coverage and sterile-to-wild male ratios [10,47,48]. Poor male competitiveness would direct attention toward rearing and release quality, whereas substantial transport-linked immigration could indicate the need for broader geographic coverage. For pgSIT, because the sterile phenotype is generated before release and is not propagated through the wild population, target site variation is not a field resistance mechanism; target and editing performance instead need to be validated during construct development and production. An IIT program would focus on compatibility, reliable female exclusion, and male performance [19,20,21], whereas Wolbachia population modification would track infection frequency, maternal transmission, host-background fitness, and pathogen blocking [22,29,36]. These examples illustrate how population genetic measurements can be translated into architecture-specific decisions; they are proposed workflows rather than field results for the example populations.

Table 1.
Evidence-to-decision map for the mosquito-control approaches examined in depth. Architecture-specific mechanisms are separated from spatial planning, which applies to every approach. Evidence limits and decision triggers are illustrative; quantitative thresholds require local prespecification and validation. Abbreviations: SIT, sterile insect technique; fsRIDL, Release of Insects carrying a Dominant Lethal; pgSIT, precision-guided sterile insect technique; IIT, incompatible insect technique; CI, cytoplasmic incompatibility; FST, fixation index.
| Approach | Measurement to collect | Evidence source | What the evidence cannot show | Illustrative decision trigger |
|---|---|---|---|---|
| Conventional SIT | Immigration, seasonal abundance, male mating success | Field and release studies [10,47,48] | Genomics alone cannot measure male mating success or field suppression. | High immigration or weak mating success: broaden coverage or revise release ratio |
| fsRIDL (female -specific dominant lethal) | Female-lethal penetrance without repressor; male survival and mating fitness; construct integrity; immigration and seasonal abundance | Laboratory studies of female-specific lethal systems [11,12]; field/release studies informing spatial deployment [10,47,48] | Construct sequence or laboratory lethality alone cannot establish female-lethal performance under field conditions, male mating success, or field suppression | Incomplete female-lethal penetrance: revise construct or rearing conditions; weak male performance: improve rearing or release ratio; high immigration: broaden coverage |
| CRISPR-based pgSIT | Editing/sterility penetrance in the production genotype; male mating fitness; immigration and seasonal abundance | Laboratory and cage tests [15,16]; field/release studies informing spatial deployment [10,47,48] |
Construct performance alone cannot establish field suppression or adequate release coverage | Incomplete sterile-phenotype penetrance: redesign guide/construct; weak male performance: improve rearing or release ratio; high immigration: broaden coverage |
| Homing gene drives | Target variants, repair outcomes, construct frequency, and fitness of escape alleles | Target-site surveys [37,40,41]; contained drive studies [91,92,93,94,95,96] | Sequence alone cannot establish functional escape; cage inheritance does not establish field spread. | functional escape allele: change target or multiplex; uncertain spread: widen monitoring |
| Threshold-dependent drives | Local introduction frequency, fitness, connectivity, seasonal demography | Spatial and threshold models [69,78,79,80,82,119,120] | Population structure or a model cannot establish a local threshold or containment. | Predicted establishment below a prespecified local range: revise site, release plan, or construct |
| Wolbachia IIT / IIT-SIT | CI profile, sex-sort leakage, male fitness, immigration | Field suppression [19,21]; dengue trial [20] | Genomes do not establish CI, female leakage, or field mating success. | Incompatibility fails or infected females appear: revise strain or sorting; high immigration: widen coverage |
| Wolbachia population modification | Host background, infection frequency, CI/compatibility, maternal transmission, virus blocking, movement | Genomic and field studies [22,29,30,36] | Genomes alone cannot show establishment or dengue protection. | Low infection frequency or stock mismatch: revise background, strain, or release area |
| Spatial planning (all approaches) | Movement between sites, immigration, seasonality, release coverage | Population structure and kinship [57,58,59,74]; spatial models [68,77,78,79] | FST or ancestry alone cannot quantify current movement, fertile introgression, or transport-linked arrivals. | Measured movement beyond planned coverage or high immigration: expand release and surveillance area |
7. From Design to Deployment: Population-based Implementation
Evidence-based deployment can be organized into three linked stages: preparation of the release stock, planning of release coverage, and evaluation of outcomes over time [35]. Release stock preparation should assess species and mating compatibility, laboratory adaptation, male or carrier fitness, and the genetic compatibility between release stocks and local populations. In a Brazilian Wolbachia population modification program, low pyrethroid resistance in the initial Ae. aegypti release stock likely contributed to failure to establish in the field after releases ceased; a subsequent release of backcrossed stock had better success in establishment due to higher insecticide resistance and higher genetic similarity to the local Ae. aegypti population [35,36]. This sequence supports evaluating release strain background and local adaptation, but it does not isolate their causal effects or establish equivalent requirements for SIT, pgSIT, or gene drives. IIT preparation additionally requires strain-specific compatibility crosses and stringent sex separation, with irradiation as a safeguard were used [19,20,21], whereas Wolbachia population modification stocks require validated maternal transmission and pathogen blocking [22].
Release planning should incorporate local vector abundance, mating structure, seasonality, species composition, and population connectivity [56,57,58,59,60,121], because these factors can influence spatial spread, population replenishment, and resistance dynamics [28,82,122]. SIT and pgSIT require sufficient releases of competitive sterile males across the intended release area, with repeated releases generally required for sustained suppression and limiting the effect of immigration potentially replenishing the target population [10,47,48]. Density-dependent larval dynamics can further complicate predictions from release ratios to subsequent adult abundance [123]. IIT similarly requires repeated releases of incompatible males together with reliable sex separation and compatibility testing [19,20,21]. Threshold-dependent drives require site-specific introduction frequencies sufficient for establishment [79,80], whereas Wolbachia population modification depends on local infection frequencies and connectivity among populations [22,29,35]. Spatial models can compare alternative release coverage across population hubs and movement corridors [61,68,77,83], but feasible deployment is also constrained by rearing capacity, sex separation, mating performance, costs, and delivery logistics [124,125].
Evaluation should use prespecified sampling before, during, and after release to distinguish intervention persistence, population replenishment, and genetic change [35]. Primary endpoints and comparators should be defined in advance and matched to the intervention architecture. For gene drives, construct frequency and functionally validated resistant haplotypes are relevant genetic outcomes. IIT evaluation should include egg hatch, wild female abundance, and surveillance for unintended establishment of the released infection, whereas Wolbachia population modification requires measurements of infection frequency, maternal transmission, and pathogen blocking. Female abundance, fertility, and vector competence provide entomological outcomes, while effects on disease incidence require appropriately designed epidemiological studies [20,22,126,127,128]. Importantly, the target of a 50% reduction in clinical malaria incidence described in [129] is a modeled outcome rather than an observed field result.
Monitoring plans should prespecify sampling locations and intervals, assays, expected ranges of outcomes, and decision rules [126,127,128]. For gene drives, an unexpected decline in construct frequency should trigger evaluation of immigration, fitness costs, construct integrity, and functionally-relevant target site resistance before release strategy or construct design is changed. For self-limiting sterile male approaches such as SIT and pgSIT, declining suppression should instead prompt evaluation of release coverage, immigration, sterile male abundance, mating competitiveness, and production quality rather than target site resistance. IIT surveillance should additionally detect infected females in the field [19,20,21], while Wolbachia population modification surveillance should track infection frequency and investigate maternal transmission, fitness, or other causes of unexpected decline [22,29,36].
Deployment also requires governance and evaluation beyond the genetic system itself. Local partners, public health authorities, regulators, and affected communities should establish expected benefits, geographic boundaries, data sharing practices, and criteria for pausing or modifying deployment [126,130]. Combining genetic or Wolbachia-based interventions with insecticides or habitat management may provide complementary control, but the effectiveness and durability of each combination require direct evaluation [131,132]. Insecticide exposure can reshape population genetic structure [133], while Wolbachia can alter reproductive compatibility and spatial patterns [29,134], making ecological context important when interventions are co-deployed. Engineered gene drives additionally require phased testing and independent risk assessment before progressively less contained evaluation [126,130].
8. Future Directions and Conclusions
Future studies should determine how well locally measured target haplotypes, Wolbachia infection dynamics, movement, mating, and demography predict intervention outcomes beyond laboratory settings. Population genomic resources remain uneven across mosquito species and geographic regions, and many identified variants lack experimentally measured effects on cleavage, repair, fitness, or resistance [135,136]. Multi-region genomic surveys should therefore be coupled with architecture-specific functional measurements, including cleavage and repair assays for sequence-targeted systems; compatibility testing for IIT; maternal transmission and pathogen-blocking assays for Wolbachia population modification; and direct measurements of movement, mating, and fitness where these parameters influence deployment. Single-cell atlases may improve tissue- and cell-type-specific target annotation [137], while repair-outcome and machine-learning models can help prioritize variants for experimental testing [112,113,114,115,116,117]. Neither, however, substitutes for functional validation in the relevant mosquito genetic background. Reporting should include negative and mixed outcomes, parameter uncertainty, and explicit limits on extrapolation across populations and sites.
A central priority is to test whether prespecified measurements and decision triggers remain informative across independent populations and deployment settings. Studies should document where forecasts succeed or fail, identify the biological or operational factors underlying discrepancies, and update models before applying release strategies to new settings. Field evidence should likewise be interpreted within the intervention and context in which it was generated. The Yogyakarta Wolbachia population modification trial [22] and the Singapore IIT-SIT trial [20], for example, provide evidence for their respective interventions in the tested settings rather than interchangeable estimates of efficacy.
Population genomics is therefore most valuable not as a stand-alone predictor of intervention success, but as part of an iterative measurement-to-decision framework that connects local genetic variation and population structure to architecture-specific functional tests, models, deployment choices, and prospective validation. Combined with community engagement and independent risk assessment, this cycle provides a testable pathway from system design to deployment and subsequent refinement (Figure 4). The framework proceeds from local baseline measurements and mechanistic testing to prespecified forecasts, independent contained evaluation, comparison of observed outcomes with predictions, and evidence-based redesign or progression.
Author Contributions
All authors contributed to the study’s conceptualization and to drafting, reviewing, and editing the manuscript.
Funding
This work was supported in part by National Institutes of Health awards R01AI151004, R21AI196522, and R01AI190001 to O.S.A., R35GM156217 to Y. L., and R01AI189916 to J.M.M. J.M.M. was also supported by funds from the Gates Foundation (INV-078535). E.H. also received support from AI007384 (HOPE T32 Training Grant).
Ethical Conduct of Research
None to declare.
Data Availability Statement
Conflicts of Interest
O.S.A. is a founder of Agragene, Inc., Synvect, Inc., and Cloak Biotechnologies with equity interest. The terms of this arrangement have been reviewed and approved by the University of California at San Diego, in accordance with its conflict of interest policies. All other authors declare no competing interests.
References
- Fu, G.L. Epidemiological patterns of mosquito-borne diseases globally. J. Mosq. Res. 2024, 14, 111–123. [Google Scholar] [CrossRef]
- Penhollow, T.M.; Torres, L.; Ferreira, M.H.; Cassol, M.; de Fraga, B.F.; Bello, J.Z.; et al. Impact of mosquito-borne diseases on global public health. Int. Phys. Med. Rehabil. J. 2021, 6, 19–20. [Google Scholar] [CrossRef]
- Bhatt, S.; Gething, P.W.; Brady, O.J.; Messina, J.P.; Farlow, A.W.; Moyes, C.L.; et al. The global distribution and burden of dengue. Nature 2013, 496, 504–507. [Google Scholar] [CrossRef] [PubMed]
- World Health Organization. World malaria report 2025: Addressing the threat of antimalarial drug resistance; World Health Organization: Geneva, 2025. [Google Scholar]
- Kondapaneni, R.; Malcolm, A.N.; Vazquez, B.M.; Zeng, E.; Chen, T.-Y.; Kosinski, K.J.; et al. Mosquito Control Priorities in Florida-Survey Results from Florida Mosquito Control Districts. Pathogens 2021, 10. [Google Scholar] [CrossRef] [PubMed]
- Hemingway, J.; Shretta, R.; Wells, T.N.C.; Bell, D.; Djimdé, A.A.; Achee, N.; et al. Tools and strategies for malaria control and elimination: What do we need to achieve a grand convergence in malaria? PLoS Biol. 2016, 14, e1002380. [Google Scholar] [CrossRef] [PubMed]
- Kumar, G.; Baharia, R.; Singh, K.; Gupta, S.K.; Joy, S.; Sharma, A.; et al. Addressing challenges in vector control: a review of current strategies and the imperative for novel tools in India’s combat against vector-borne diseases. BMJ Public Health 2024, 2, e000342. [Google Scholar] [CrossRef] [PubMed]
- Beier, J.C.; Keating, J.; Githure, J.I.; Macdonald, M.B.; Impoinvil, D.E.; Novak, R.J. Integrated vector management for malaria control. Malar. J. 2008, 7, S4. [Google Scholar] [CrossRef] [PubMed]
- Knipling, E.F. Possibilities of insect control or eradication through the use of sexually sterile Males1. J. Econ. Entomol. 1955, 48, 459–462. [Google Scholar] [CrossRef]
- Dyck, V.A.; Hendrichs, J.; Robinson, A.S. Sterile insect technique: Principles and practice in area-wide integrated pest management; Springer: New York, NY, 2005. [Google Scholar]
- Labbé, G.M.; Scaife, S.; Morgan, S.A.; Curtis, Z.H.; Alphey, L. Female-specific flightless (fsRIDL) phenotype for control of Aedes albopictus. PLoS Negl. Trop. Dis. 2012, 6, e1724. [Google Scholar] [CrossRef] [PubMed]
- Spinner, S.A.M.; Barnes, Z.H.; Puinean, A.M.; Gray, P.; Dafa’alla, T.; Phillips, C.E.; et al. New self-sexing Aedes aegypti strain eliminates barriers to scalable and sustainable vector control for governments and communities in dengue-prone environments. Front Bioeng. Biotechnol. 2022, 10, 975786. [Google Scholar] [CrossRef] [PubMed]
- Smidler, A.L.; Akbari, O.S. CRISPR technologies for the control and study of malaria-transmitting anopheline mosquitoes. Parasit. Vectors 2025, 18, 252. [Google Scholar] [CrossRef] [PubMed]
- Kandul, N.P.; Liu, J.; Sanchez, C. H.M.; Wu, S.L.; Marshall, J.M.; Akbari, O.S. Transforming insect population control with precision guided sterile males with demonstration in flies. Nat. Commun. 2019, 10, 84. [Google Scholar] [CrossRef] [PubMed]
- Li, M.; Yang, T.; Bui, M.; Gamez, S.; Wise, T.; Kandul, N.P.; et al. Suppressing mosquito populations with precision guided sterile males. Nat. Commun. 2021, 12, 5374. [Google Scholar] [CrossRef] [PubMed]
- Apte, R.A.; Smidler, A.L.; Pai, J.J.; Chow, M.L.; Chen, S.; Mondal, A.; et al. Eliminating malaria vectors with precision-guided sterile males. Proc. Natl. Acad. Sci. U S A 2024, 121, e2312456121. [Google Scholar] [CrossRef] [PubMed]
- Esvelt, K.M.; Smidler, A.L.; Catteruccia, F.; Church, G.M. Concerning RNA-guided gene drives for the alteration of wild populations. Elife 2014, 3, e03401. [Google Scholar] [CrossRef] [PubMed]
- Champer, J.; Buchman, A.; Akbari, O.S. Cheating evolution: engineering gene drives to manipulate the fate of wild populations. Nat. Rev. Genet. 2016, 17, 146–159. [Google Scholar] [CrossRef] [PubMed]
- Zeng, Q.; She, L.; Yuan, H.; Luo, Y.; Wang, R.; Mao, W.; et al. A standalone incompatible insect technique enables mosquito suppression in the urban subtropics. Commun. Biol. 2022, 5, 1419. [Google Scholar] [CrossRef] [PubMed]
- Lim, J.T.; Chong, C.-S.; Chang, C.-C.; Mailepessov, D.; Dickens, B.; Lai, Y.L.; et al. Dengue suppression by male Wolbachia-infected mosquitoes. N Engl. J. Med. 2026, 394, 1175–1183. [Google Scholar] [CrossRef] [PubMed]
- Zheng, X.; Zhang, D.; Li, Y.; Yang, C.; Wu, Y.; Liang, X.; et al. Incompatible and sterile insect techniques combined eliminate mosquitoes. Nature 2019, 572, 56–61. [Google Scholar] [CrossRef] [PubMed]
- Utarini, A.; Indriani, C.; Ahmad, R.A.; Tantowijoyo, W.; Arguni, E.; Ansari, M.R.; et al. Efficacy of Wolbachia-infected mosquito deployments for the control of dengue. N Engl. J. Med. 2021, 384, 2177–2186. [Google Scholar] [CrossRef] [PubMed]
- Collier, T.C.; Lee, Y.; Mathias, D.K.; López Del Amo, V. CRISPR-Cas9 and Cas12a target site richness reflects genomic diversity in natural populations of Anopheles gambiae and Aedes aegypti mosquitoes. BMC Genom. 2024, 25, 700. [Google Scholar] [CrossRef] [PubMed]
- Schmidt, H.; Collier, T.C.; Hanemaaijer, M.J.; Houston, P.D.; Lee, Y.; Lanzaro, G.C. Abundance of conserved CRISPR-Cas9 target sites within the highly polymorphic genomes of Anopheles and Aedes mosquitoes. Nat. Commun. 2020, 11, 1425. [Google Scholar] [CrossRef] [PubMed]
- Schmidt, H.; Lee, Y.; Collier, T.C.; Hanemaaijer, M.J.; Kirstein, O.D.; Ouledi, A.; et al. Transcontinental dispersal of occurred from West African origin via serial founder events. Commun. Biol. 2019, 2, 473. [Google Scholar] [CrossRef] [PubMed]
- Campos, M.; Hanemaaijer, M.; Gripkey, H.; Collier, T.C.; Lee, Y.; Cornel, A.J.; et al. The origin of island populations of the African malaria mosquito, Anopheles coluzzii. Commun. Biol. 2021, 4, 630. [Google Scholar] [CrossRef] [PubMed]
- Zentner, G.E.; Wade, M.J. The promise and peril of CRISPR gene drives: Genetic variation and inbreeding may impede the propagation of gene drives based on the CRISPR genome editing technology: Genetic variation and inbreeding may impede the propagation of gene drives based on the CRISPR genome editing technology. Bioessays 2017, 39, 1700109. [Google Scholar] [CrossRef] [PubMed]
- Unckless, R.L.; Clark, A.G.; Messer, P.W. Evolution of resistance against CRISPR/Cas9 gene drive. Genetics 2017, 205, 827–841. [Google Scholar] [CrossRef] [PubMed]
- Schmidt, T.L.; Filipović, I.; Hoffmann, A.A.; Rašić, G. Fine-scale landscape genomics helps explain the slow spatial spread of Wolbachia through the Aedes aegypti population in Cairns, Australia. Heredity (Edinb) 2018, 120, 386–395. [Google Scholar] [CrossRef] [PubMed]
- Beeton, N.J.; Wilkins, A.; Ickowicz, A.; Hayes, K.R.; Hosack, G.R. Spatial modelling for population replacement of mosquito vectors at continental scale. PLoS Comput Biol. 2022, 18, e1009526. [Google Scholar] [CrossRef] [PubMed]
- Gomulkiewicz, R.; Thies, M.L.; Bull, J.J. Evading resistance to gene drives. Genetics 2021, 217, iyaa040. [Google Scholar] [CrossRef] [PubMed]
- Gloria-Soria, A.; Soghigian, J.; Kellner, D.; Powell, J.R. Genetic diversity of laboratory strains and implications for research: The case of Aedes aegypti. PLoS Negl. Trop. Dis. 2019, 13, e0007930. [Google Scholar] [CrossRef] [PubMed]
- Alphey, L. Genetic control of mosquitoes. Annu Rev. Entomol. 2014, 59, 205–224. [Google Scholar] [CrossRef] [PubMed]
- Raban, R.; James, A.A.; Akbari, O.S. Advances in CRISPR gene drives for mosquito population control. Curr. Opin. Microbiol. 2026, 90, 102712. [Google Scholar] [CrossRef] [PubMed]
- Schmidt, T.L.; Endersby-Harshman, N.M.; Hoffmann, A.A. Improving mosquito control strategies with population genomics. Trends Parasitol. 2021, 37, 907–921. [Google Scholar] [CrossRef] [PubMed]
- Garcia, G.; de, A.; Sylvestre, G.; Aguiar, R.; da Costa, G.B.; Martins, A.J.; Lima, J.B.P.; et al. Matching the genetics of released and local Aedes aegypti populations is critical to assure Wolbachia invasion. PLoS Negl. Trop. Dis. 2019, 13, e0007023. [Google Scholar] [CrossRef] [PubMed]
- Schmidt, H.; Collier, T.C.; Hanemaaijer, M.J.; Houston, P.D.; Lee, Y.; Lanzaro, G.C. Abundance of conserved CRISPR-Cas9 target sites within the highly polymorphic genomes of Anopheles and Aedes mosquitoes. Nat. Commun. 2020, 11, 1425. [Google Scholar] [CrossRef] [PubMed]
- Weeraratne, T.C.; Surendran, S.N.; Walton, C.; Karunaratne, S.H.P.P. Genetic diversity and population structure of malaria vector mosquitoes Anopheles subpictus, Anopheles peditaeniatus, and Anopheles vagus in five districts of Sri Lanka. Malar. J. 2018, 17, 271. [Google Scholar] [CrossRef] [PubMed]
- Ahebwa, A.; Hii, J.; Neoh, K.-B.; Chareonviriyaphap, T. Aedes aegypti and Aedes albopictus (Diptera: Culicidae) ecology, biology, behaviour, and implications on arbovirus transmission in Thailand: Review. One Health 2023, 16, 100555. [Google Scholar] [CrossRef] [PubMed]
- Collier, T.C.; Lee, Y.; Mathias, D.K.; López Del Amo, V. CRISPR-Cas9 and Cas12a target site richness reflects genomic diversity in natural populations of Anopheles gambiae and Aedes aegypti mosquitoes. BMC Genom. 2024, 25, 700. [Google Scholar] [CrossRef] [PubMed]
- Pescod, P.; Bevivino, G.; Anthousi, A.; Shelton, R.; Shepherd, J.; Lombardo, F.; et al. Measuring the impact of genetic heterogeneity and chromosomal inversions on the efficacy of CRISPR-Cas9 gene drives in different strains of Anopheles gambiae. CRISPR J. 2023, 6, 419–429. [Google Scholar] [CrossRef] [PubMed]
- Pescod, P.; Bevivino, G.; Anthousi, A.; Shepherd, J.; Shelton, R.; Lombardo, F.; et al. Homing gene drives can transfer rapidly between Anopheles gambiae strains with minimal carryover of flanking sequences. Nat. Commun. 2024, 15, 6846. [Google Scholar] [CrossRef] [PubMed]
- Facchinelli, L.; Valerio, L.; Ramsey, J.M.; Gould, F.; Walsh, R.K.; Bond, G.; et al. Field cage studies and progressive evaluation of genetically-engineered mosquitoes. PLoS Negl. Trop. Dis. 2013, 7, e2001. [Google Scholar] [CrossRef] [PubMed]
- Aldersley, A.; Pongsiri, A.; Bunmee, K.; Kijchalao, U.; Chittham, W.; Fansiri, T.; et al. Too “sexy” for the field? Paired measures of laboratory and semi-field performance highlight variability in the apparent mating fitness of Aedes aegypti transgenic strains. Parasit. Vectors 2019, 12, 357. [Google Scholar] [CrossRef] [PubMed]
- Drury, D.W.; Dapper, A.L.; Siniard, D.J.; Zentner, G.E.; Wade, M.J. CRISPR/Cas9 gene drives in genetically variable and nonrandomly mating wild populations. Sci. Adv. 2017, 3, e1601910. [Google Scholar] [CrossRef] [PubMed]
- Burt, A. Site-specific selfish genes as tools for the control and genetic engineering of natural populations. Proc. Biol. Sci. 2003, 270, 921–928. [Google Scholar] [CrossRef] [PubMed]
- Alphey, L.; Benedict, M.; Bellini, R.; Clark, G.G.; Dame, D.A.; Service, M.W.; et al. Sterile-insect methods for control of mosquito-borne diseases: an analysis. Vector Borne Zoonotic Dis. 2010, 10, 295–311. [Google Scholar] [CrossRef] [PubMed]
- Bourtzis, K.; Vreysen, M.J.B. Sterile insect technique (SIT) and its applications. Insects 2021, 12, 638. [Google Scholar] [CrossRef] [PubMed]
- Li, M.; Kandul, N.P.; Sun, R.; Yang, T.; Benetta, E.D.; Brogan, D.J.; et al. Targeting sex determination to suppress mosquito populations. Elife 2024, 12, RP90199. [Google Scholar] [CrossRef] [PubMed]
- Kandul, N.P.; Liu, J.; Buchman, A.; Shriner, I.C.; Corder, R.M.; Warsinger-Pepe, N.; et al. Precision guided sterile males suppress populations of an invasive crop pest. GEN Biotechnol. 2022, 1, 372–385. [Google Scholar] [CrossRef]
- Anderson, M.A.E.; Gonzalez, E.; Edgington, M.P.; Ang, J.X.D.; Purusothaman, D.-K.; Shackleford, L.; et al. A multiplexed, confinable CRISPR/Cas9 gene drive can propagate in caged Aedes aegypti populations. Nat. Commun. 2024, 15, 729. [Google Scholar] [CrossRef] [PubMed]
- Larrosa-Godall, M.; Shackleford, L.; Edgington, M.P.; Leftwich, P.T.; Luk, J.C.Y.; Southworth, J.; et al. Integrating multiplexing into confineable gene drives effectively overrides resistance in Anopheles stephensi. Nat. Commun. 2026, 17, 6844. [Google Scholar] [CrossRef] [PubMed]
- Lacroix, R.; McKemey, A.R.; Raduan, N.; Kwee Wee, L.; Hong Ming, W.; Guat Ney, T.; et al. Open field release of genetically engineered sterile male Aedes aegypti in Malaysia. PLoS ONE 2012, 7, e42771. [Google Scholar] [CrossRef] [PubMed]
- Beeton, N.J.; Wilkins, A.; Ickowicz, A.; Hayes, K.R.; Hosack, G.R. Spatial modelling for population replacement of mosquito vectors at continental scale. PLoS Comput Biol. 2022, 18, e1009526. [Google Scholar] [CrossRef] [PubMed]
- Lee, Y.; Schmidt, H.; Collier, T.C.; Conner, W.R.; Hanemaaijer, M.J.; Slatkin, M.; et al. Genome-wide divergence among invasive populations of Aedes aegypti in California. BMC Genom. 2019, 20, 204. [Google Scholar] [CrossRef] [PubMed]
- Taylor, C.; Touré, Y.T.; Carnahan, J.; Norris, D.E.; Dolo, G.; Traoré, S.F.; et al. Gene flow among populations of the malaria vector, Anopheles gambiae, in Mali, West Africa. Genetics 2001, 157, 743–750. [Google Scholar] [CrossRef] [PubMed]
- Sumitha, M.K.; Kalimuthu, M.; Kumar, M.S.; Paramasivan, R.; Kumar, N.P.; Sunish, I.P.; et al. Genetic differentiation among Aedes aegypti populations from different eco-geographical zones of India. PLoS Negl. Trop. Dis. 2023, 17, e0011486. [Google Scholar] [CrossRef] [PubMed]
- Hlaing, T.; Tun-Lin, W.; Somboon, P.; Socheat, D.; Setha, T.; Min, S.; et al. Spatial genetic structure of Aedes aegypti mosquitoes in mainland Southeast Asia: Genetic structure ofAedes aegypti. Evol. Appl. 2010, 3, 319–339. [Google Scholar] [CrossRef] [PubMed]
- Failloux, A.-B.; Vazeille, M.; Rodhain, F. Geographic genetic variation in populations of the dengue virus vector Aedes aegypti. J. Mol. Evol. 2002, 55, 653–663. [Google Scholar] [CrossRef] [PubMed]
- Besansky, N.J.; Lehmann, T.; Fahey, G.T.; Fontenille, D.; Braack, L.E.; Hawley, W.A. Patterns of mitochondrial variation within and between African malaria vectors, Anopheles gambiae and An. arabiensis, suggest extensive gene flow. Genetics 1997, 147, 1817–1828. [Google Scholar] [CrossRef] [PubMed]
- Hemme, R.R.; Thomas, C.L.; Chadee, D.D.; Severson, D.W. Influence of urban landscapes on population dynamics in a short-distance migrant mosquito: evidence for the dengue vector Aedes aegypti. PLoS Negl. Trop. Dis. 2010, 4, e634. [Google Scholar] [CrossRef] [PubMed]
- Lee, Y.; Marsden, C.D.; Norris, L.C.; Collier, T.C.; Main, B.J.; Fofana, A.; et al. Spatiotemporal dynamics of gene flow and hybrid fitness between the M and S forms of the malaria mosquito, Anopheles gambiae. Proc. Natl. Acad. Sci. U S A 2013, 110, 19854–19859. [Google Scholar] [CrossRef] [PubMed]
- Norris, L.C.; Main, B.J.; Lee, Y.; Collier, T.C.; Fofana, A.; Cornel, A.J.; et al. Adaptive introgression in an African malaria mosquito coincident with the increased usage of insecticide-treated bed nets. Proc. Natl. Acad. Sci. U S A 2015, 112, 815–820. [Google Scholar] [CrossRef] [PubMed]
- Main, B.J.; Lee, Y.; Collier, T.C.; Norris, L.C.; Brisco, K.; Fofana, A.; et al. Complex genome evolution in Anopheles coluzzii associated with increased insecticide usage in Mali. Mol. Ecol. 2015, 24, 5145–5157. [Google Scholar] [CrossRef] [PubMed]
- Hanemaaijer, M.J.; Collier, T.C.; Chang, A.; Shott, C.C.; Houston, P.D.; Schmidt, H.; et al. The fate of genes that cross species boundaries after a major hybridization event in a natural mosquito population. Mol. Ecol. 2018, 27, 4978–4990. [Google Scholar] [CrossRef] [PubMed]
- Willoughby, J.R.; McKenzie, B.A.; Ahn, J.; Steury, T.D.; Lepzcyk, C.A.; Zohdy, S. Assessing and managing the risk of Aedes mosquito introductions via the global maritime trade network. PLoS Negl. Trop. Dis. 2024, 18, e0012110. [Google Scholar] [CrossRef] [PubMed]
- Ahn, J.; Sinka, M.; Irish, S.; Zohdy, S. Modeling marine cargo traffic to identify countries in Africa with greatest risk of invasion by Anopheles stephensi. Sci. Rep. 2023, 13, 876. [Google Scholar] [CrossRef] [PubMed]
- Eckhoff, P.A.; Wenger, E.A.; Godfray, H.C.J.; Burt, A. Impact of mosquito gene drive on malaria elimination in a computational model with explicit spatial and temporal dynamics. Proc. Natl. Acad. Sci. U S A 2017, 114, E255–E264. [Google Scholar] [CrossRef] [PubMed]
- Noble, C.; Adlam, B.; Church, G.M.; Esvelt, K.M.; Nowak, M.A. Current CRISPR gene drive systems are likely to be highly invasive in wild populations. Elife 2018, 7, e33423. [Google Scholar] [CrossRef] [PubMed]
- Champer, J.; Wen, Z.; Luthra, A.; Reeves, R.; Chung, J.; Liu, C.; et al. CRISPR gene drive efficiency and resistance rate is highly heritable with no common genetic loci of large effect. Genetics 2019, 212, 333–341. [Google Scholar] [CrossRef] [PubMed]
- Ickowicz, A.; Foster, S.D.; Hosack, G.R.; Hayes, K.R. Predicting the spread and persistence of genetically modified dominant sterile male mosquitoes. Parasit. Vectors 2021, 14, 480. [Google Scholar] [CrossRef] [PubMed]
- Marshall, J.M.; Yang, S.; Bennett, J.B.; Filipović, I.; Rašić, G. Spatial close-kin mark-recapture methods to estimate dispersal parameters and barrier strength for mosquitoes. PLoS Comput Biol. 2025, 21, e1013713. [Google Scholar] [CrossRef] [PubMed]
- Sánchez, C. H.M.; Wu, S.L.; Henry, J.M.; Guerra, C.A.; Galick, D.S.; Garcia, G.A.; et al. Mosquito dispersal in context. PLoS Complex Syst. 2026, 3, e0000108. [Google Scholar] [CrossRef]
- Jasper, M.; Schmidt, T.L.; Ahmad, N.W.; Sinkins, S.P.; Hoffmann, A.A. A genomic approach to inferring kinship reveals limited intergenerational dispersal in the yellow fever mosquito. Mol. Ecol. Resour. 2019, 19, 1254–1264. [Google Scholar] [CrossRef] [PubMed]
- Barton, N.H.; Turelli, M. Spatial waves of advance with bistable dynamics: cytoplasmic and genetic analogues of Allee effects. Am. Nat. 2011, 178, E48–75. [Google Scholar] [CrossRef] [PubMed]
- Hanski, I. Metapopulation dynamics. Nature 1998, 396, 41–49. [Google Scholar] [CrossRef]
- Wu, S.L.; Bennett, J.B.; Sánchez, C. H.M.; Dolgert, A.J.; León, T.M.; Marshall, J.M. MGDrivE 2: A simulation framework for gene drive systems incorporating seasonality and epidemiological dynamics. PLoS Comput Biol. 2021, 17, e1009030. [Google Scholar] [CrossRef] [PubMed]
- Champer, J.; Zhao, J.; Champer, S.E.; Liu, J.; Messer, P.W. Population dynamics of underdominance gene drive systems in continuous space. ACS Synth. Biol. 2020, 9, 779–792. [Google Scholar] [CrossRef] [PubMed]
- Sánchez, C. H.M.; Bennett, J.B.; Wu, S.L.; Rašić, G.; Akbari, O.S.; Marshall, J.M. Modeling confinement and reversibility of threshold-dependent gene drive systems in spatially-explicit Aedes aegypti populations. BMC Biol. 2020, 18, 50. [Google Scholar] [CrossRef] [PubMed]
- Kim, I.K.; Messer, P.W. Predicting the invasiveness of threshold-dependent gene drives. bioRxivorg. bioRxiv 2025. [Google Scholar] [CrossRef] [PubMed]
- Li, M.; Yang, T.; Kandul, N.P.; Bui, M.; Gamez, S.; Raban, R.; et al. Development of a confinable gene drive system in the human disease vector Aedes aegypti. Elife 2020, 9, e51701. [Google Scholar] [CrossRef] [PubMed]
- Marshall, J.M.; Hay, B.A. Confinement of gene drive systems to local populations: a comparative analysis. J. Theor. Biol. 2012, 294, 153–171. [Google Scholar] [CrossRef] [PubMed]
- North, A.; Burt, A.; Godfray, H.C.J. Modelling the spatial spread of a homing endonuclease gene in a mosquito population. J. Appl. Ecol. 2013, 50, 1216–1225. [Google Scholar] [CrossRef] [PubMed]
- Price, T.A.R.; Windbichler, N.; Unckless, R.L.; Sutter, A.; Runge, J.-N.; Ross, P.A.; et al. Resistance to natural and synthetic gene drive systems. J. Evol. Biol. 2020, 33, 1345–1360. [Google Scholar] [CrossRef] [PubMed]
- Romanowski, J.S.; Myles, K.M.; Adelman, Z.N. Microhomology-mediated end joining is the predominant form of DNA repair in the mosquito Aedes aegypti with implications for gene editing, gene drive, and transgene removal. Nucleic Acids Res. 2026, 54, gkaf1532. [Google Scholar] [CrossRef] [PubMed]
- Hou, S.; Chen, J.; Feng, R.; Xu, X.; Liang, N.; Champer, J. A homing rescue gene drive with multiplexed gRNAs reaches high frequency in cage populations but generates functional resistance. J. Genet Genom. 2024, 51, 836–843. [Google Scholar] [CrossRef] [PubMed]
- Khatri, B.S.; Burt, A. A theory of resistance to multiplexed gene drive demonstrates the significant role of weakly deleterious natural genetic variation. Proc. Natl. Acad. Sci. U S A 2022, 119, e2200567119. [Google Scholar] [CrossRef] [PubMed]
- Hammond, A.; Galizi, R.; Kyrou, K.; Simoni, A.; Siniscalchi, C.; Katsanos, D.; et al. A CRISPR-Cas9 gene drive system targeting female reproduction in the malaria mosquito vector Anopheles gambiae. Nat. Biotechnol. 2016, 34, 78–83. [Google Scholar] [CrossRef] [PubMed]
- Oberhofer, G.; Ivy, T.; Hay, B.A. Behavior of homing endonuclease gene drives targeting genes required for viability or female fertility with multiplexed guide RNAs. Proc. Natl. Acad. Sci. U S A 2018, 115, E9343–E9352. [Google Scholar] [CrossRef] [PubMed]
- Hammond, A.M.; Kyrou, K.; Bruttini, M.; North, A.; Galizi, R.; Karlsson, X.; et al. The creation and selection of mutations resistant to a gene drive over multiple generations in the malaria mosquito. PLoS Genet. 2017, 13, e1007039. [Google Scholar] [CrossRef] [PubMed]
- Gantz, V.M.; Jasinskiene, N.; Tatarenkova, O.; Fazekas, A.; Macias, V.M.; Bier, E.; et al. Highly efficient Cas9-mediated gene drive for population modification of the malaria vector mosquito Anopheles stephensi. Proc. Natl. Acad. Sci. U S A 2015, 112, E6736–43. [Google Scholar] [CrossRef] [PubMed]
- Champer, J.; Reeves, R.; Oh, S.Y.; Liu, C.; Liu, J.; Clark, A.G.; et al. Novel CRISPR/Cas9 gene drive constructs reveal insights into mechanisms of resistance allele formation and drive efficiency in genetically diverse populations. PLoS Genet. 2017, 13, e1006796. [Google Scholar] [CrossRef] [PubMed]
- Carrami, E.M.; Eckermann, K.N.; Ahmed, H.M.M.; Sánchez, C. H.M.; Dippel, S.; Marshall, J.M.; et al. Consequences of resistance evolution in a Cas9-based sex conversion-suppression gene drive for insect pest management. Proc. Natl. Acad. Sci. U S A 2018, 115, 6189–6194. [Google Scholar] [CrossRef] [PubMed]
- Hammond, A.; Pollegioni, P.; Persampieri, T.; North, A.; Minuz, R.; Trusso, A.; et al. Gene-drive suppression of mosquito populations in large cages as a bridge between lab and field. Nat. Commun. 2021, 12, 4589. [Google Scholar] [CrossRef] [PubMed]
- Carballar-Lejarazú, R.; Dong, Y.; Pham, T.B.; Tushar, T.; Stillinger, D.; Nguyen, D.N.; et al. Long-term stability and performance of Cas9/guide RNA-based gene drives in anopheline mosquitoes. Proc. Natl. Acad. Sci. U S A 2026, 123, e2605739123. [Google Scholar] [CrossRef] [PubMed]
- Morianou, I.; Phillimore, L.; Khatri, B.S.; Marston, L.; Gribble, M.; Burt, A.; et al. Engineering resilient gene drives for sustainable malaria control by predicting, testing and overcoming target site resistance in Anopheles gambiae. PLoS Biol. 2026, 24, e3003879. [Google Scholar] [CrossRef] [PubMed]
- Champer, J.; Lee, E.; Yang, E.; Liu, C.; Clark, A.G.; Messer, P.W. A toxin-antidote CRISPR gene drive system for regional population modification. Nat. Commun. 2020, 11, 1082. [Google Scholar] [CrossRef] [PubMed]
- Faber, N.R.; McFarlane, G.R.; Gaynor, R.C.; Pocrnic, I.; Whitelaw, C.B.A.; Gorjanc, G. Novel combination of CRISPR-based gene drives eliminates resistance and localises spread. Sci. Rep. 2021, 11, 3719. [Google Scholar] [CrossRef] [PubMed]
- Marshall, J.M.; Buchman, A.; Sánchez, C. H.M.; Akbari, O.S. Overcoming evolved resistance to population-suppressing homing-based gene drives. Sci. Rep. 2017, 7, 3776. [Google Scholar] [CrossRef] [PubMed]
- Kandul, N.P.; Liu, J.; Bennett, J.B.; Marshall, J.M.; Akbari, O.S. A confinable home-and-rescue gene drive for population modification. Elife 2021, 10. [Google Scholar] [CrossRef] [PubMed]
- Oberhofer, G.; Ivy, T.; Hay, B.A. Cleave and Rescue, a novel selfish genetic element and general strategy for gene drive. Proc. Natl. Acad. Sci. U S A 2019, 116, 6250–6259. [Google Scholar] [CrossRef] [PubMed]
- Finney, M.; Romanowski, J.; Adelman, Z.N. Strategies to improve homology-based repair outcomes following CRISPR-based gene editing in mosquitoes: lessons in how to keep any repair disruptions local. Virol. J. 2022, 19, 128. [Google Scholar] [CrossRef] [PubMed]
- Champer, J.; Yang, E.; Lee, E.; Liu, J.; Clark, A.G.; Messer, P.W. A CRISPR homing gene drive targeting a haplolethal gene removes resistance alleles and successfully spreads through a cage population. Proc. Natl. Acad. Sci. U S A 2020, 117, 24377–24383. [Google Scholar] [CrossRef] [PubMed]
- Zhu, J.; Chen, J.; Liu, Y.; Xu, X.; Champer, J. Population suppression with dominant female-lethal alleles is boosted by homing gene drive. BMC Biol. 2024, 22, 201. [Google Scholar] [CrossRef] [PubMed]
- Carballar-Lejarazú, R.; Tushar, T.; Pham, T.B.; James, A.A. Cas9-mediated maternal effect and derived resistance alleles in a gene-drive strain of the African malaria vector mosquito, Anopheles gambiae. Genetics 2022, 221, iyac055. [Google Scholar] [CrossRef] [PubMed]
- Kosinski, K.; Lee, Y.; Romero-Weaver, A.; Chen, T.-Y.; Collier, T.; Wang, X.; et al. TWO NOVEL SINGLE NUCLEOTIDE POLYMORPHISMS IN THE VOLTAGE-GATED SODIUM CHANNEL GENE IDENTIFIED IN AEDES AEGYPTI MOSQUITOES FROM FLORIDA. J. FL Mosq. Control Assoc. 2022, 69. [Google Scholar] [CrossRef]
- Zheng, X.; Zhang, D.; Li, Y.; Yang, C.; Wu, Y.; Liang, X.; et al. Incompatible and sterile insect techniques combined eliminate mosquitoes. Nature 2019, 572, 56–61. [Google Scholar] [CrossRef] [PubMed]
- Zhang, D.; Zheng, X.; Xi, Z.; Bourtzis, K.; Gilles, J.R.L. Combining the sterile insect technique with the incompatible insect technique: I-impact of wolbachia infection on the fitness of triple- and double-infected strains of Aedes albopictus. PLoS ONE 2015, 10, e0121126. [Google Scholar] [CrossRef] [PubMed]
- Lutrat, C.; Giesbrecht, D.; Marois, E.; Whyard, S.; Baldet, T.; Bouyer, J. Sex Sorting for Pest Control: It’s Raining Men! Trends Parasitol. 2019, 35, 649–662. [Google Scholar] [CrossRef] [PubMed]
- Ross, P.A.; Wiwatanaratanabutr, I.; Axford, J.K.; White, V.L.; Endersby-Harshman, N.M.; Hoffmann, A.A. Wolbachia Infections in Aedes aegypti Differ Markedly in Their Response to Cyclical Heat Stress. PLoS Pathog. 2017, 13, e1006006. [Google Scholar] [CrossRef] [PubMed]
- Ross, P.A.; Axford, J.K.; Yang, Q.; Staunton, K.M.; Ritchie, S.A.; Richardson, K.M.; et al. Heatwaves cause fluctuations in wMel Wolbachia densities and frequencies in Aedes aegypti. PLoS Negl. Trop. Dis. 2020, 14, e0007958. [Google Scholar] [CrossRef] [PubMed]
- Allen, F.; Crepaldi, L.; Alsinet, C.; Strong, A.J.; Kleshchevnikov, V.; De Angeli, P.; et al. Predicting the mutations generated by repair of Cas9-induced double-strand breaks. Nat. Biotechnol. 2018, 37, 64–72. [Google Scholar] [CrossRef] [PubMed]
- Shen, M.W.; Arbab, M.; Hsu, J.Y.; Worstell, D.; Culbertson, S.J.; Krabbe, O.; et al. Predictable and precise template-free CRISPR editing of pathogenic variants. Nature 2018, 563, 646–651. [Google Scholar] [CrossRef] [PubMed]
- Chen, W.; McKenna, A.; Schreiber, J.; Haeussler, M.; Yin, Y.; Agarwal, V.; et al. Massively parallel profiling and predictive modeling of the outcomes of CRISPR/Cas9-mediated double-strand break repair. Nucleic Acids Res. 2019, 47, 7989–8003. [Google Scholar] [CrossRef] [PubMed]
- Xiang, X.; Corsi, G.I.; Anthon, C.; Qu, K.; Pan, X.; Liang, X.; et al. Enhancing CRISPR-Cas9 gRNA efficiency prediction by data integration and deep learning. Nat. Commun. 2021, 12, 3238. [Google Scholar] [CrossRef] [PubMed]
- Liu, X.; Wang, S.; Ai, D. Predicting CRISPR/Cas9 repair outcomes by Attention-based deep learning framework. Cells 2022, 11, 1847. [Google Scholar] [CrossRef] [PubMed]
- Kim, M.-G.; Go, M.-J.; Kang, S.-H.; Jeong, S.-H.; Lim, K. Revolutionizing CRISPR technology with artificial intelligence. Exp. Mol. Med. 2025, 57, 1419–1431. [Google Scholar] [CrossRef] [PubMed]
- Rašić, G.; Marshall, J.M. Integrating mosquito genomics into simulation modeling: opportunities for better-informed biocontrol. Curr. Opin. Insect Sci. 2026, 73, 101456. [Google Scholar] [CrossRef] [PubMed]
- Gendron, W.A.C.; Smidler, A.L.; Li, M.; Wise, T.; Sánchez, C. H.M.; Bennett, J.B.; et al. Split suppression gene drives reveal a design tradeoff between inheritance and reproductive load in. bioRxiv 2026. [Google Scholar] [CrossRef] [PubMed]
- Verkuijl, S.A.N.; Gonzalez, E.; Li, M.; Ang, J.X.D.; Kandul, N.P.; Anderson, M.A.E.; et al. A CRISPR endonuclease gene drive reveals distinct mechanisms of inheritance bias. Nat. Commun. 2022, 13, 7145. [Google Scholar] [CrossRef] [PubMed]
- Dhole, S.; Lloyd, A.L.; Gould, F. Gene drive dynamics in natural populations: The importance of density dependence, space, and sex. Annu Rev. Ecol. Evol. Syst. 2020, 51, 505–531. [Google Scholar] [CrossRef] [PubMed]
- Dhole, S.; Lloyd, A.L.; Gould, F. Tethered homing gene drives: A new design for spatially restricted population replacement and suppression. Evol. Appl. 2019, 12, 1688–1702. [Google Scholar] [CrossRef] [PubMed]
- Legros, M.; Lloyd, A.L.; Huang, Y.; Gould, F. Density-dependent intraspecific competition in the larval stage of Aedes aegypti (Diptera: Culicidae): revisiting the current paradigm. J. Med. Entomol. 2009, 46, 409–419. [Google Scholar] [CrossRef] [PubMed]
- Gendron, W.A.C.; Raban, R.; Mondal, A.; M Sánchez, C. H.; Smidler, A.L.; Zilberman, D.; et al. Evaluating the cost of malaria elimination by Anopheles gambiae precision guided SIT in the Upper River region, The Gambia. PLoS Glob. Public Health 2025, 5, e0004903. [Google Scholar] [CrossRef] [PubMed]
- Gendron, W.A.C.; Raban, R.; Mondal, A.; Sánchez, C. H.M.; Zilberman, D.; Ilboudo, P.G.C.; et al. Estimated cost and operational structure of pgSIT malaria vector control programs in selected West African countries. Sci. Afr. 2025, 29, None. [Google Scholar] [CrossRef] [PubMed]
- James, S.; Collins, F.H.; Welkhoff, P.A.; Emerson, C.; Godfray, H.C.J.; Gottlieb, M.; et al. Pathway to deployment of gene drive mosquitoes as a potential biocontrol tool for elimination of malaria in sub-Saharan Africa: Recommendations of a scientific working group. Am. J. Trop. Med. Hyg. 2018, 98, 1–49. [Google Scholar] [CrossRef] [PubMed]
- Okumu, F.O.; Finda, M.; Odero, J.O.; Aboagye-Antwi, F.; Adeogun, A.; Atta-Obeng, C.; et al. Adapting Africa’s vector surveillance systems to monitor gene-drive mosquitoes in malaria control. Trends Parasitol. 2026, 42, 463–493. [Google Scholar] [CrossRef]
- Rašić, G.; Lobo, N.F.; Jeffrey Gutiérrez, E.H.; Sánchez, C. H.M.; Marshall, J.M. Monitoring needs for gene drive mosquito projects: Lessons from vector control field trials and invasive species. Front Genet. 2021, 12, 780327. [Google Scholar] [CrossRef] [PubMed]
- Mondal, A.; Sánchez, C. H.M.; Marshall, J.M. A model-informed target product profile for population modification gene drives for malaria control. PLoS Glob. Public Health 2025, 5, e0005026. [Google Scholar] [CrossRef] [PubMed]
- Devos, Y.; Mumford, J.D.; Bonsall, M.B.; Glandorf, D.C.M.; Quemada, H.D. Risk management recommendations for environmental releases of gene drive modified insects. Biotechnol. Adv. 2022, 54, 107807. [Google Scholar] [CrossRef] [PubMed]
- Abbasi, E. Innovative approaches to vector control: integrating genomic, biological, and chemical strategies. Ann. Med. Surg. (Lond) 2025, 87, 5003–5011. [Google Scholar] [CrossRef] [PubMed]
- Tourapi, C.; Tsioutis, C. Circular Policy: A new approach to vector and vector-borne diseases’ management in line with the global vector control response (2017-2030). Trop. Med. Infect. Dis. 2022, 7, 125. [Google Scholar] [CrossRef] [PubMed]
- Hancock, P.A.; Hendriks, C.J.M.; Tangena, J.-A.; Gibson, H.; Hemingway, J.; Coleman, M.; et al. Mapping trends in insecticide resistance phenotypes in African malaria vectors. PLoS Biol. 2020, 18, e3000633. [Google Scholar] [CrossRef] [PubMed]
- O’Neill, S.L.; Ryan, P.A.; Turley, A.P.; Wilson, G.; Retzki, K.; Iturbe-Ormaetxe, I.; et al. Scaled deployment of Wolbachia to protect the community from dengue and other Aedes transmitted arboviruses. Gates Open Res. 2019, 2, 36. Available online: https://pmc.ncbi.nlm.nih.gov/articles/PMC6305154/. [CrossRef] [PubMed]
- Ruzzante, L.; Reijnders, M.J.; Waterhouse, R.M. Of genes and genomes: mosquito evolution and diversity. Trends Parasitol. 2019, 35, 32–51. Available online: https://www.cell.com/trends/parasitology/fulltext/S1471-4922(18)30220-4. [CrossRef] [PubMed]
- Neafsey, D.E.; Taylor, A.R.; MacInnis, B.L. Advances and opportunities in malaria population genomics. Nat. Rev. Genet. 2021, 22, 502–517. Available online: https://www.nature.com/articles/s41576-021-00349-5. [CrossRef] [PubMed]
- Nayyar, N.; Arya, S.K.; Harrison, D.A.; Palli, S.R. Recent advances and applications of single-cell sequencing in insects. Curr. Opin. Insect Sci. 2026, 73, 101455. [Google Scholar] [CrossRef] [PubMed]
- Website. [CrossRef]
Figure 1.
Population genetic measurements guide choices across population suppression approaches (SIT, fsRIDL, pgSIT, and Wolbachia IIT) and population replacement approaches (homing drives and Wolbachia population modification). Target haplotypes, connectivity, and demography inform design; genetic and entomological monitoring test the forecast and guide revision. Field surveys, contained tests, and models provide distinct evidence. Arrows show conceptual decisions, not demonstrated field efficacy.
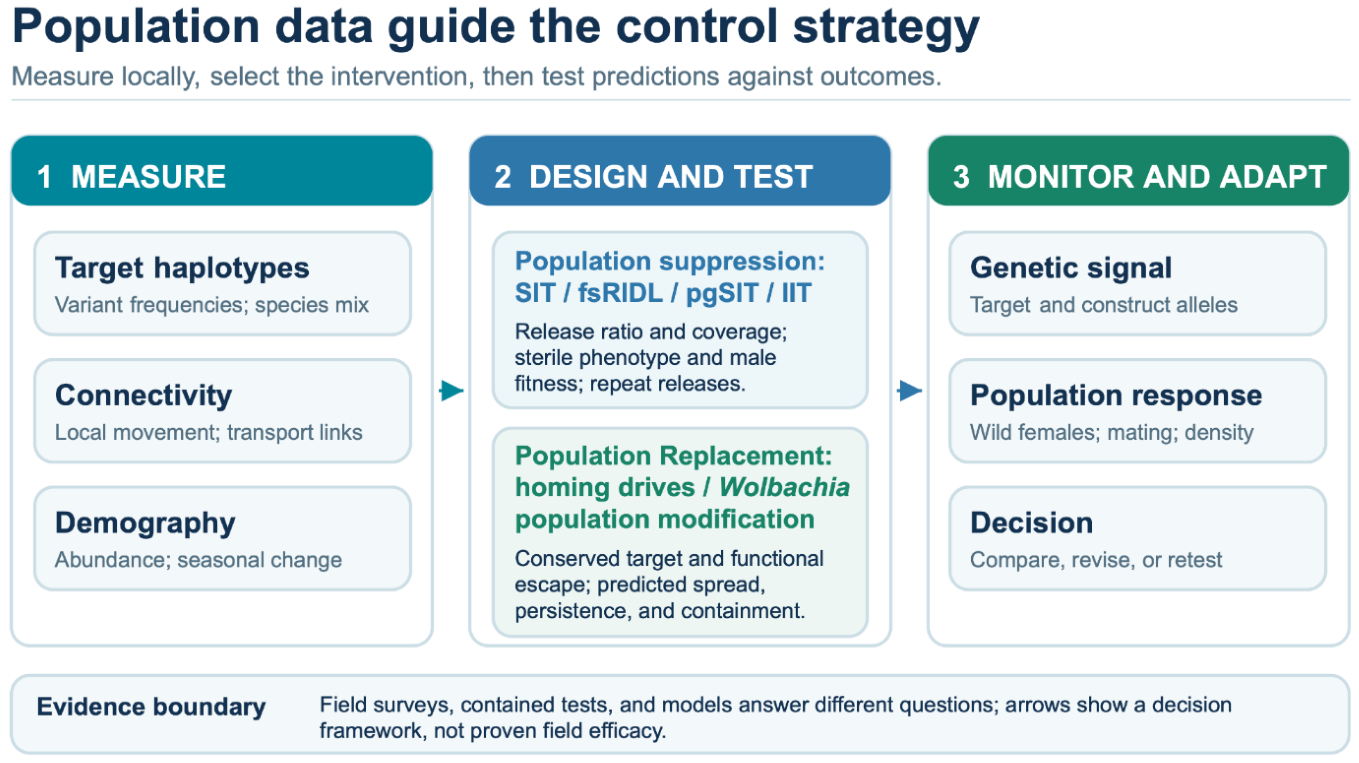
Figure 2.
Conceptual population structure and connectivity in Anopheles and Aedes. Panel A uses blue for An. gambiae, orange for An. coluzzii, and purple for An. arabiensis to show varying species composition across sites; cross-species gene flow requires fertile crosses. Panel B uses green for Ae. aegypti in a neighborhood network. Solid arrows mark movement links to test, and dashed arrows mark possible transport links. Dots and arrows are schematic, not measured frequencies or migration rates. Local sampling must determine the real network [56,57,58,59,60,61].
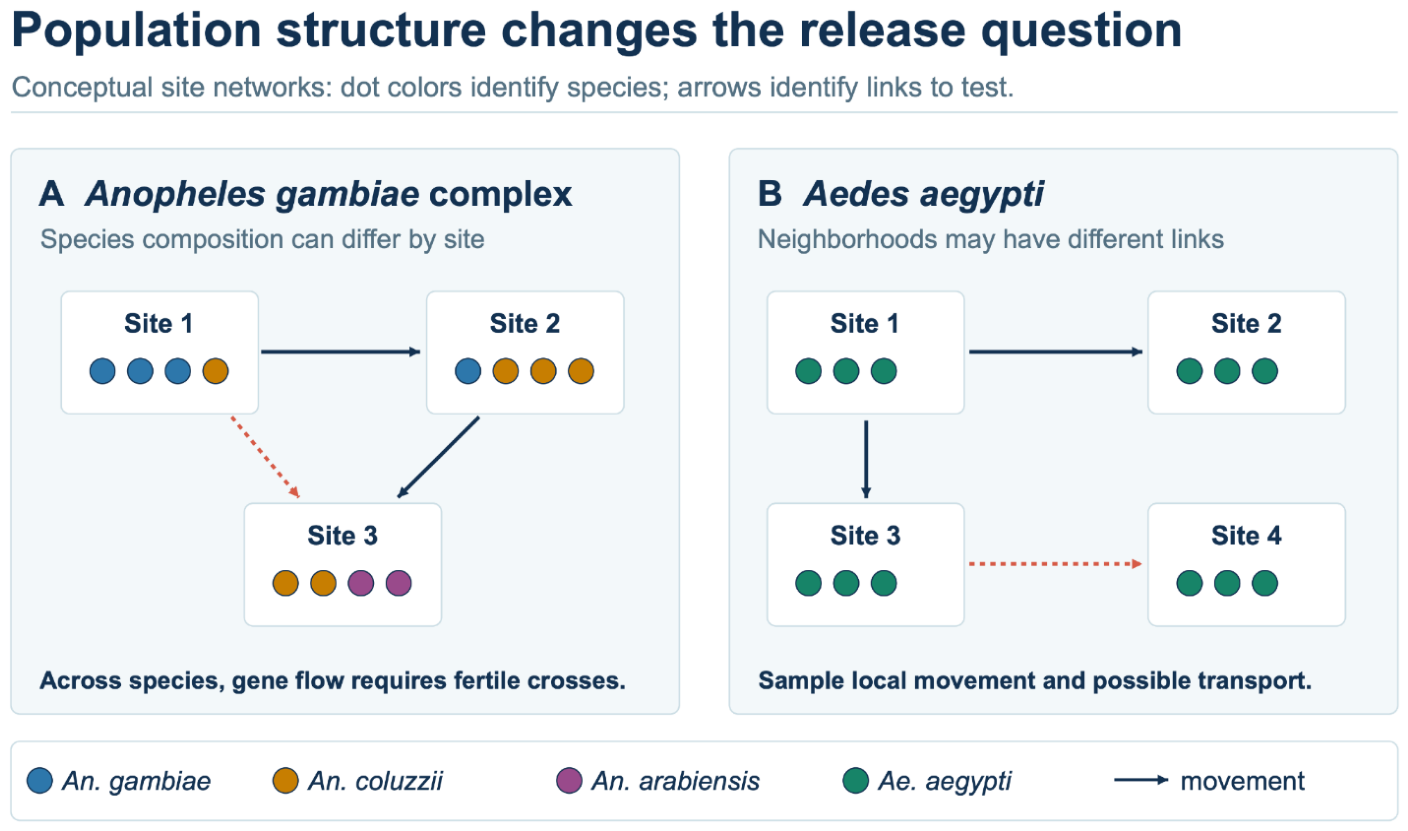
Figure 3.
Homing-drive resistance classification and illustrative consequences for suppression and population-replacement drives.
Figure 3.
Homing-drive resistance classification and illustrative consequences for suppression and population-replacement drives.
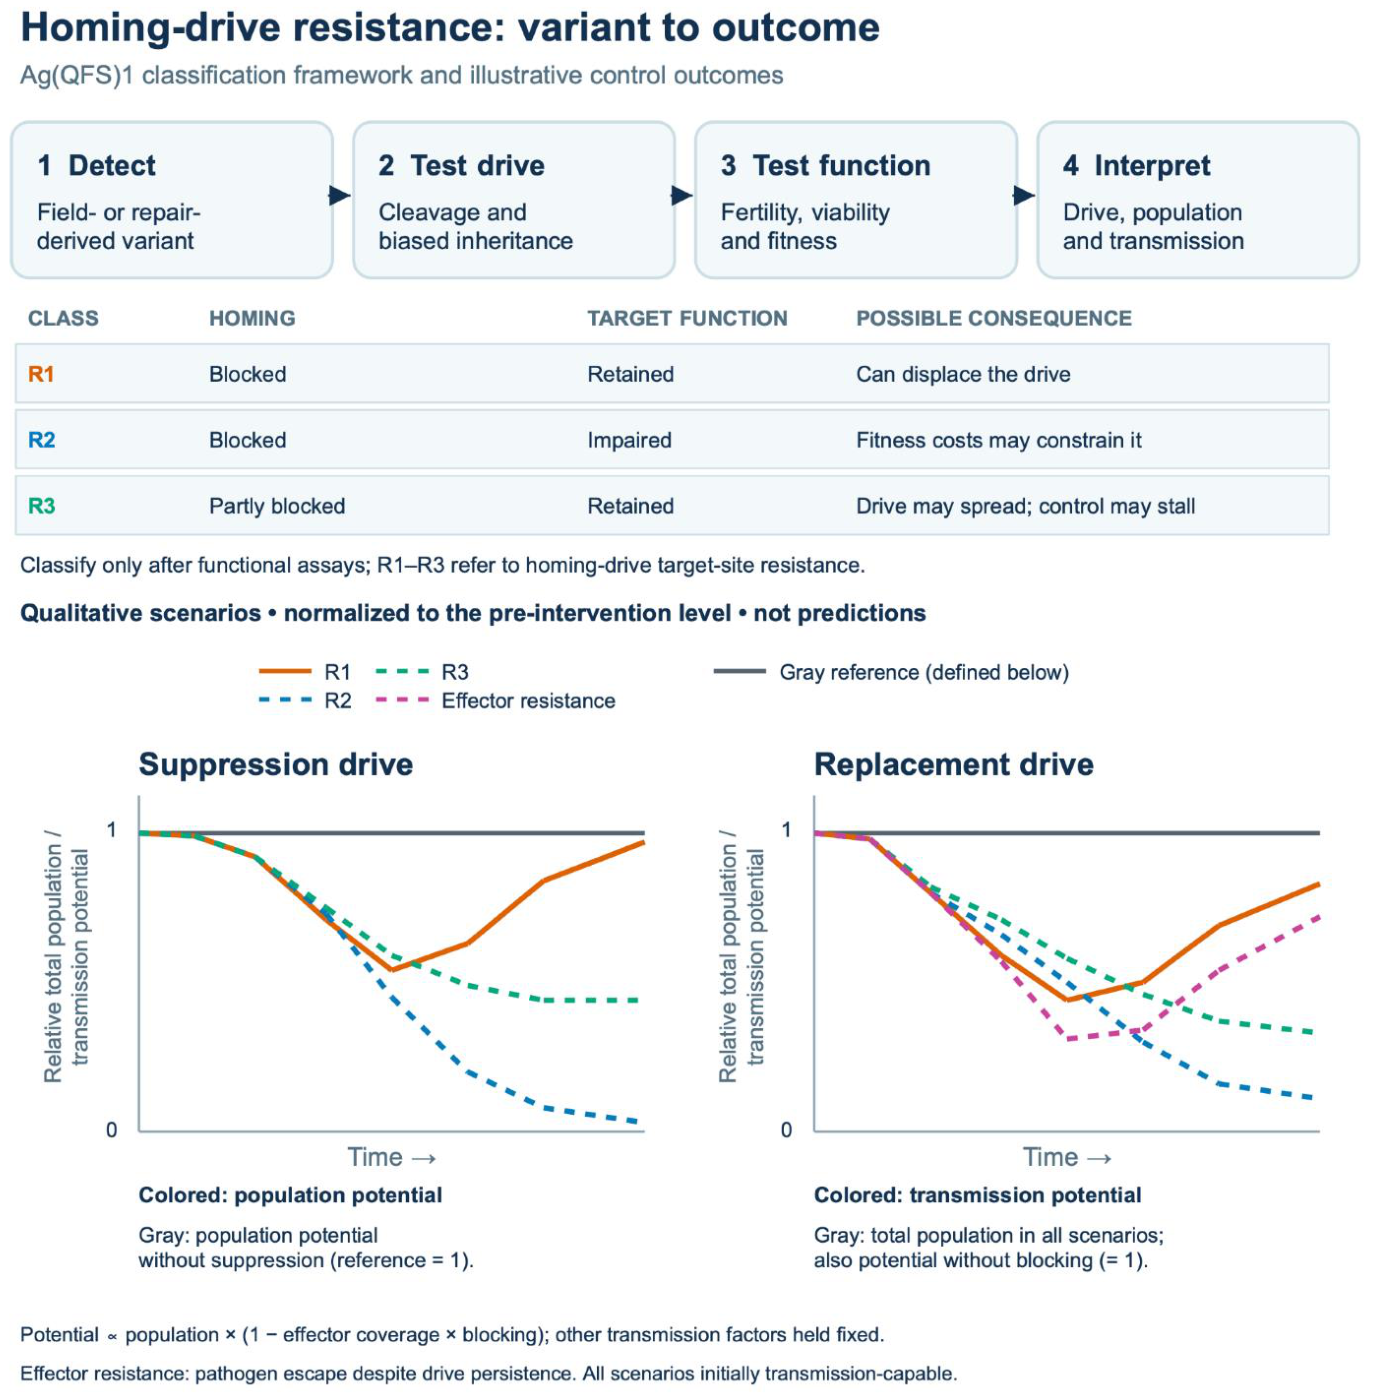
Figure 4.
Prospective validation pathway for population-informed control. Local genomic, movement, and demographic measurements plus functional assays define forecasts, endpoints, and failure modes before testing. Independent contained tests compare genetic and entomological outcomes with predictions; discrepancies prompt redesign or new measurements. Epidemiological impact requires separate field evidence, and progression needs local engagement, risk assessment, and regulatory review [126,127,128,130].
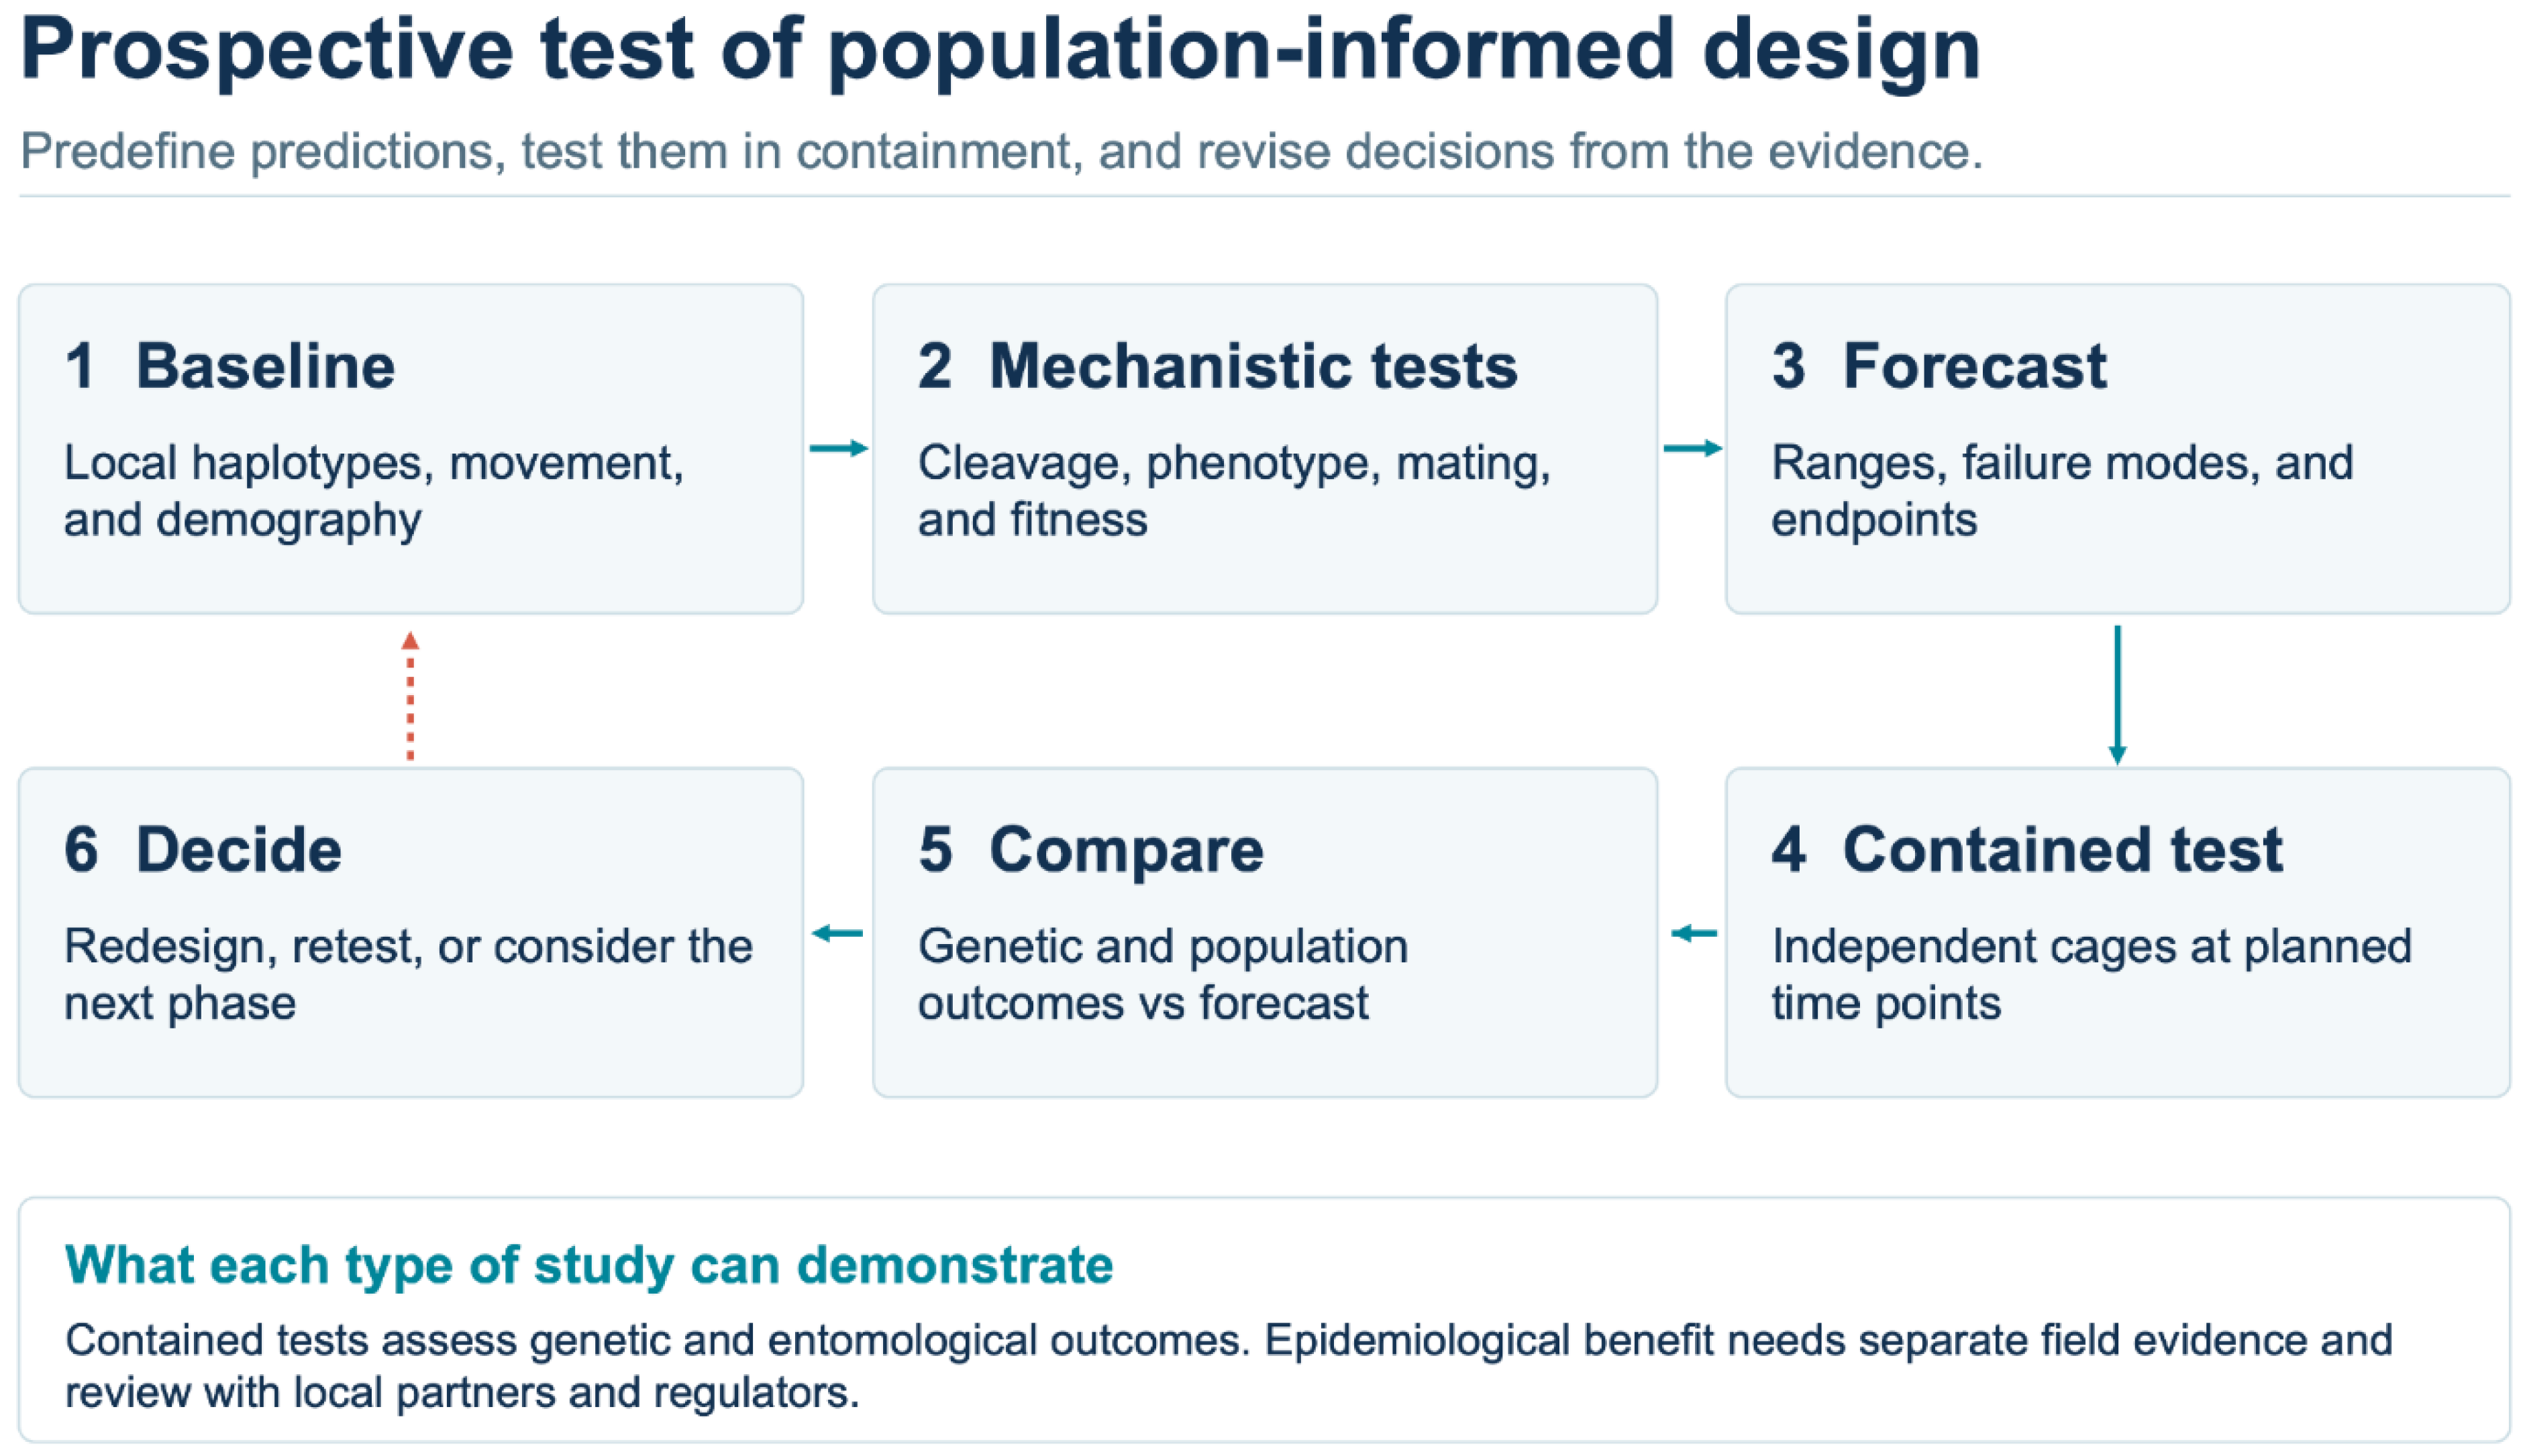
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (http://creativecommons.org/licenses/by/4.0/).
Copyright: This open access article is published under a Creative Commons CC BY 4.0 license, which permit the free download, distribution, and reuse, provided that the author and preprint are cited in any reuse.