

Submitted:
23 September 2026
Posted:
24 September 2026
You are already at the latest version
Abstract
The apparent yield of “full” recombinant adeno-associated virus (rAAV) capsids is a key manufacturing metric. Yet it can substantially overstate the fraction of particles that contain a single intact, transduction-competent genome. I propose that a stochastic occupancy ceiling constrains authentic full-capsid yield, a limit that bulk production assays often neglect. Single-particle and single-molecule studies show that the density- and mass-defined “full” population is heterogeneous, containing truncated, rearranged, and multi-DNA species alongside intact genomes, implying stochastic rather than deterministic loading. In the resulting occupancy model, the fraction of capsids containing exactly one intact genome is p(1) = m·e^(-m), where m is the mean number of genome-equivalents per capsid. This fraction maximizes at m = 1, imposing an e^(-1) ≈ 36.8% ceiling on authentic single-genome capsids in any independent, batch-loading regime. Increasing genome supply beyond this point cannot increase authentically-full yield; instead, it converts empty capsids into physically full but functionally defective multi-occupancy particles that co-purify with authentic capsids and evade density-, size-, and bulk-DNA assays. Reports of high-percentage full-capsid fractions should therefore be interpreted with caution unless supported by sequence-level evidence of single intact genomes. The model also identifies the only route to exceed this ceiling: occupancy-dependent protection, represented by the relative second-capture parameter, σ, combined with decoupled, low-instantaneous-MOE (Multiplicity-Of-Encapsidation) loading. This framework reframes rAAV filling from a separations problem to a quantitative loading-control problem. Single-molecule sequencing of the full-capsid density fraction generated under varied genome supply would be the decisive analytical test of the hypothesis.
Keywords:
adeno-associated virus
; AAV manufacturing
; empty and full capsids
; defective-interfering particles
; stochastic packaging
; Poisson statistics
; multi-channel insertion
; single-molecule sequencing
; gene therapy
1. Introduction
Recombinant adeno-associated virus (rAAV) is the leading platform for in vivo gene therapy, with several approved products and a large clinical pipeline (Kuzmin et al. 2021, Wang et al. 2019). A persistent manufacturing challenge is the low and variable proportion of capsids that carry the intended therapeutic genome. The conventional paradigm distinguishes “empty” capsids, which lack a genome, from “full” capsids, which are assumed to contain one, and frames process optimization as maximizing the full-to-empty ratio. Empty capsids are considered liabilities because they may be immunogenic, compete for receptor binding and intracellular trafficking, and dilute the effective dose. Accordingly, substantial effort has focused on increasing the full-length fraction and on separating full from empty capsids by density or charge (Penaud-Budloo et al. 2018).
This paper offers a different interpretation. I argue that the empty/full dichotomy overlooks a third, analytically hidden class of particles; that the authentic full-capsid fraction, i.e., capsids carrying a single intact, ITR-flanked, and expression-competent genome, is bound by a statistical ceiling that conventional process optimization cannot overcome, and that this ceiling lies well below the values commonly reported for purified preparations.
The argument has three components. First, published single-particle studies already show that the “full” population is heterogeneous in genomic content. Second, this heterogeneity is incompatible with strictly deterministic, single-genome packaging and instead implies stochastic loading. Third, a simple stochastic-occupancy model predicts an upper bound of about 36.8% for authentic single-genome capsids in any independent single-batch production run and indicates that increasing the genome-to-capsid ratio converts empty capsids into defective particles rather than authentic, transduction-competent ones.
The central point of this argument is the “stealth-defective” particle: a physically full capsid, and therefore indistinguishable from an authentic capsid by density or mass, but whose packaged DNA fragments do not constitute a single intact genome. The quantitative ceiling proposed here does not depend on any specific molecular mechanism of encapsidation; rather, it follows purely from loading stochastics. Nevertheless, Section 4.1 proposes a plausible controlled molecular mechanism for exceeding the stochastic limit on genuinely full (authentic) AAV particles and explains its rationale.
The empirical premise for this interpretation is not merely that some particles are full by physical criteria, but that the full-density fraction is compositionally heterogeneous. Single-particle and single-molecule analyses show that capsids classified as full by density or mass can contain incomplete genome fragments, rearranged molecules, or other noncanonical DNA species rather than a single intact vector genome.
Single-molecule sequencing of the purified full-density rAAV fraction (≈1.39–1.45 g/cm³) revealed that many recovered molecules were less than half the genome length. In the foundational study, 5′-containing fragments shorter than half the genome length accounted for roughly half of all 5′ ends sequenced from that fraction (Kapranov et al. 2012). Subsequent long-read and next-generation sequencing studies of AAV populations have identified snapback, truncated, rearranged, and other subgenomic species co-resident with canonical genomes. Some apparently homogeneous preparations contained well under half full-length species (Lecomte et al. 2015, Tai et al. 2018, Zhang et al. 2022). Charge-detection and native mass spectrometry of intact particles further resolve species consistent with partial genomes, heterogeneous non-vector DNA, and particles whose mass indicates more than one genome-equivalent of packaged nucleic acid (Nakatsuka et al. 2024, Pierson et al. 2016).
If packaging produced exactly one intact genome in every genome-containing capsid, as a strictly single-genome, processive “fill-to-capacity-then-stop” mechanism would imply, the full-density fraction should be approximately homogeneous in its single-intact-genome content. It is not. The observed heterogeneity therefore provides direct evidence that genome loading departs from strict single-genome determinism and includes a stochastic component.
2. Materials and Methods
2.1. The Stochastic-Occupancy Model
I modeled genome loading as a stochastic process across a population of preformed capsids. The mean number of genome-equivalents loaded per capsid, denoted by “m,” is referred to here as the multiplicity of encapsidation (MOE). Under the minimal assumption that loading events are independent across the available genome supply, the number of genome-equivalents in an individual capsid follows a Poisson distribution:
giving three operationally relevant occupancy classes:
p(k) = m^(k) · e^(−m) / k!
Empty Fraction: p(0) = e^(-m)
Authentic Single-Genome Fraction: p(1) = m · e^(-m)
Multi-occupancy (stealth-defective) Fraction: p(≥2) = 1 – e^(-m)(1+m)
Here, the “loading event” refers to incorporating one genome-equivalent of DNA into a capsid.
2.2. Model Development and Computational Methods
I first derived the hypotheses and governing mathematical relationships by hand using Poisson statistics and polynomial series expansions; I presented and defended portions of this work in the cited poster. During manuscript preparation, Claude Opus 4.8 (Anthropic) was used, under the author’s direction, to improve notation, presentation, and computational implementation. Specifically, it helped translate my hand-derived relationships into compact symbolic form, define and formalize the protection coefficient σ as the relative second-capture rate, extend the discrete loading model to the continuous fully staged limit using ordinary differential equations, and compute and plot the model across parameter space. Independently, I checked the AI-assisted derivations, figures, and conclusions against my original hand derivations and verified that the stated assumptions support the manuscript's claims. Full model details are provided in Appendices B and C.
3. Results
3.1. The Ceiling
Differentiating p(1) = m·e^(−m): d/dm = e^(−m)(1 − m) = 0 at m = 1, so
p(1)|_(m=1) = e^(−1) ≈ 0.3679 ≈ 36.8%.
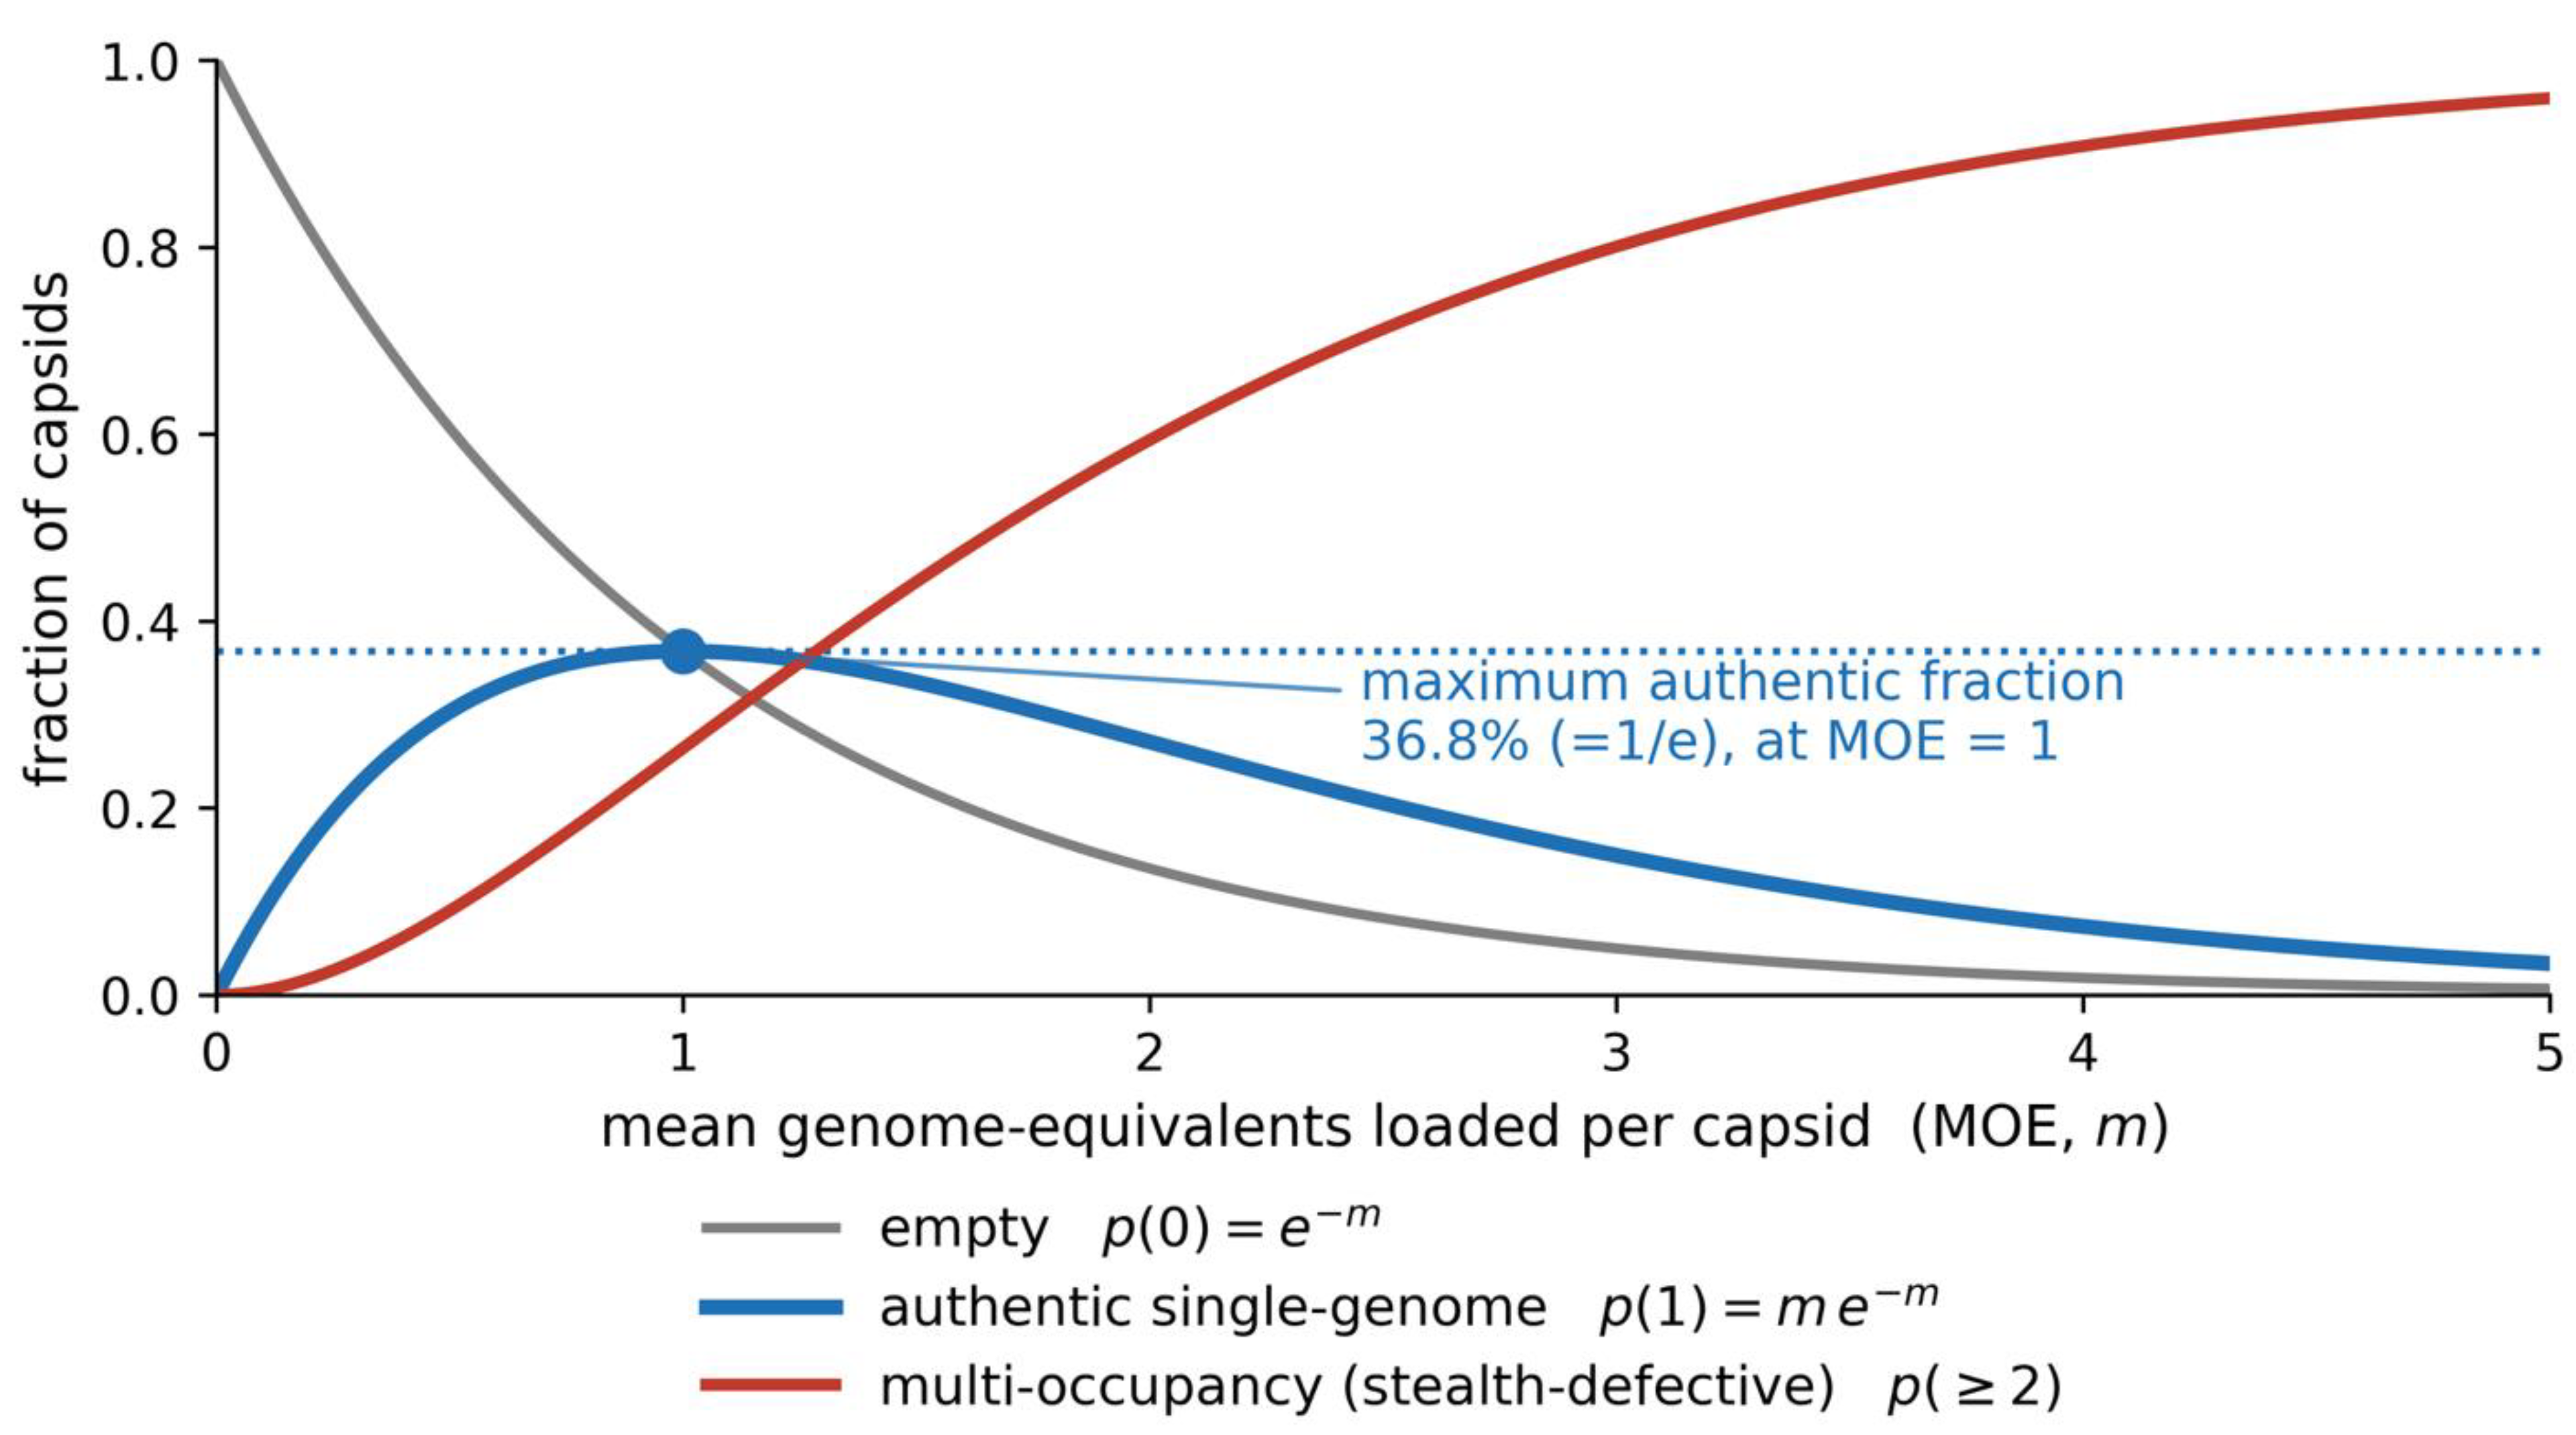
At the optimum, the model predicts three coexisting classes: approximately 36.8% empty capsids, 36.8% authentic single-genome capsids, and 26.4% multi-occupancy particles. The maximum authentic fraction is therefore much lower than the high “percent full” values often reported for purified preparations. This 36.8% maximum is the central quantitative claim of Section 3.1.
3.2. Why Increasing Genome Supply Can Reduce Authentic Yield
Section 3.2 explains the consequences of moving away from the optimum. Increasing genome supply reduces the empty-capsid fraction, but the effects differ before and after m = 1. When m is below 1, added genomes primarily increase the authentic single-genome class, although many capsids remain empty. Once m exceeds 1, additional loading events increasingly generate multi-occupancy particles. The apparent full fraction can therefore continue to rise even as the authentic single-genome fraction falls. In this regime, adding more genomes mainly converts empty capsids into physically full but functionally defective stealth-defective particles. Strategies that increase genome availability without increasing capsid number are therefore predicted to enrich defective full particles rather than authentic single-genome vectors.
Figure 1.
Poisson occupancy distribution versus mean genome-equivalents per capsid (MOE): empty e^(−m), authentic m·e^(−m), multi-occupancy 1 − e^(−m)(1+m); for m = 0→5, the authentic maximum (36.8%) occurs at m = 1.
Figure 1.
Poisson occupancy distribution versus mean genome-equivalents per capsid (MOE): empty e^(−m), authentic m·e^(−m), multi-occupancy 1 − e^(−m)(1+m); for m = 0→5, the authentic maximum (36.8%) occurs at m = 1.
3.3. Scope and Relaxation of the Stochastic Bound
The bound represents the theoretical maximum for authentic genome encapsidation under stochastic, independent loading. If packaging were strictly processive and limited to one genome per capsid, no such ceiling would exist, because genome-containing capsids would be expected to be nearly uniformly single-genome. The heterogeneity described in the Introduction therefore provides the empirical basis for rejecting a strict deterministic model.
The Poisson formulation further assumes that loading events are independent and evenly distributed; the most consequential extension of this view concerns whether successive loading events on the same capsid remain independent. If a capsid that has captured one genome becomes less likely to capture another—for example, through interior volume exclusion, conformational sealing, or viral Rep motor dissociation—then the single-occupancy state is protected, and the authentic fraction can exceed 1/e. Section 3.4 formalizes this possibility with a single parameter, σ, defined as the relative rate at which an already-loaded capsid captures an additional genome. In this framework, the 36.8% ceiling applies when σ = 1 (no protection). Finally, the model counts genome-equivalents rather than intact genomes; the distinction between single occupancy and a single intact genome is an empirical factor addressed in Section 4.3.
3.4. Exceeding the Ceiling Through Occupancy-Dependent Protection
The authentic capsid ceiling corresponds to the σ = 1 limit, where σ is the rate at which a single-occupancy capsid captures an additional genome relative to the rate at which an empty capsid captures its first genome. Authentic yield can exceed 1/e only if σ < 1, indicating that a capsid already containing one genome is protected from further capture. To model decoupled loading explicitly, we held the capsid number fixed while introducing genomes over R rounds at a per-empty-capsid dose “α” (the instantaneous MOE), with single-occupancy capsids capturing additional genomes at the relative rate σ. This setup has clear implications that naturally reduce the anticipated advantages of staged (incremental) loading.
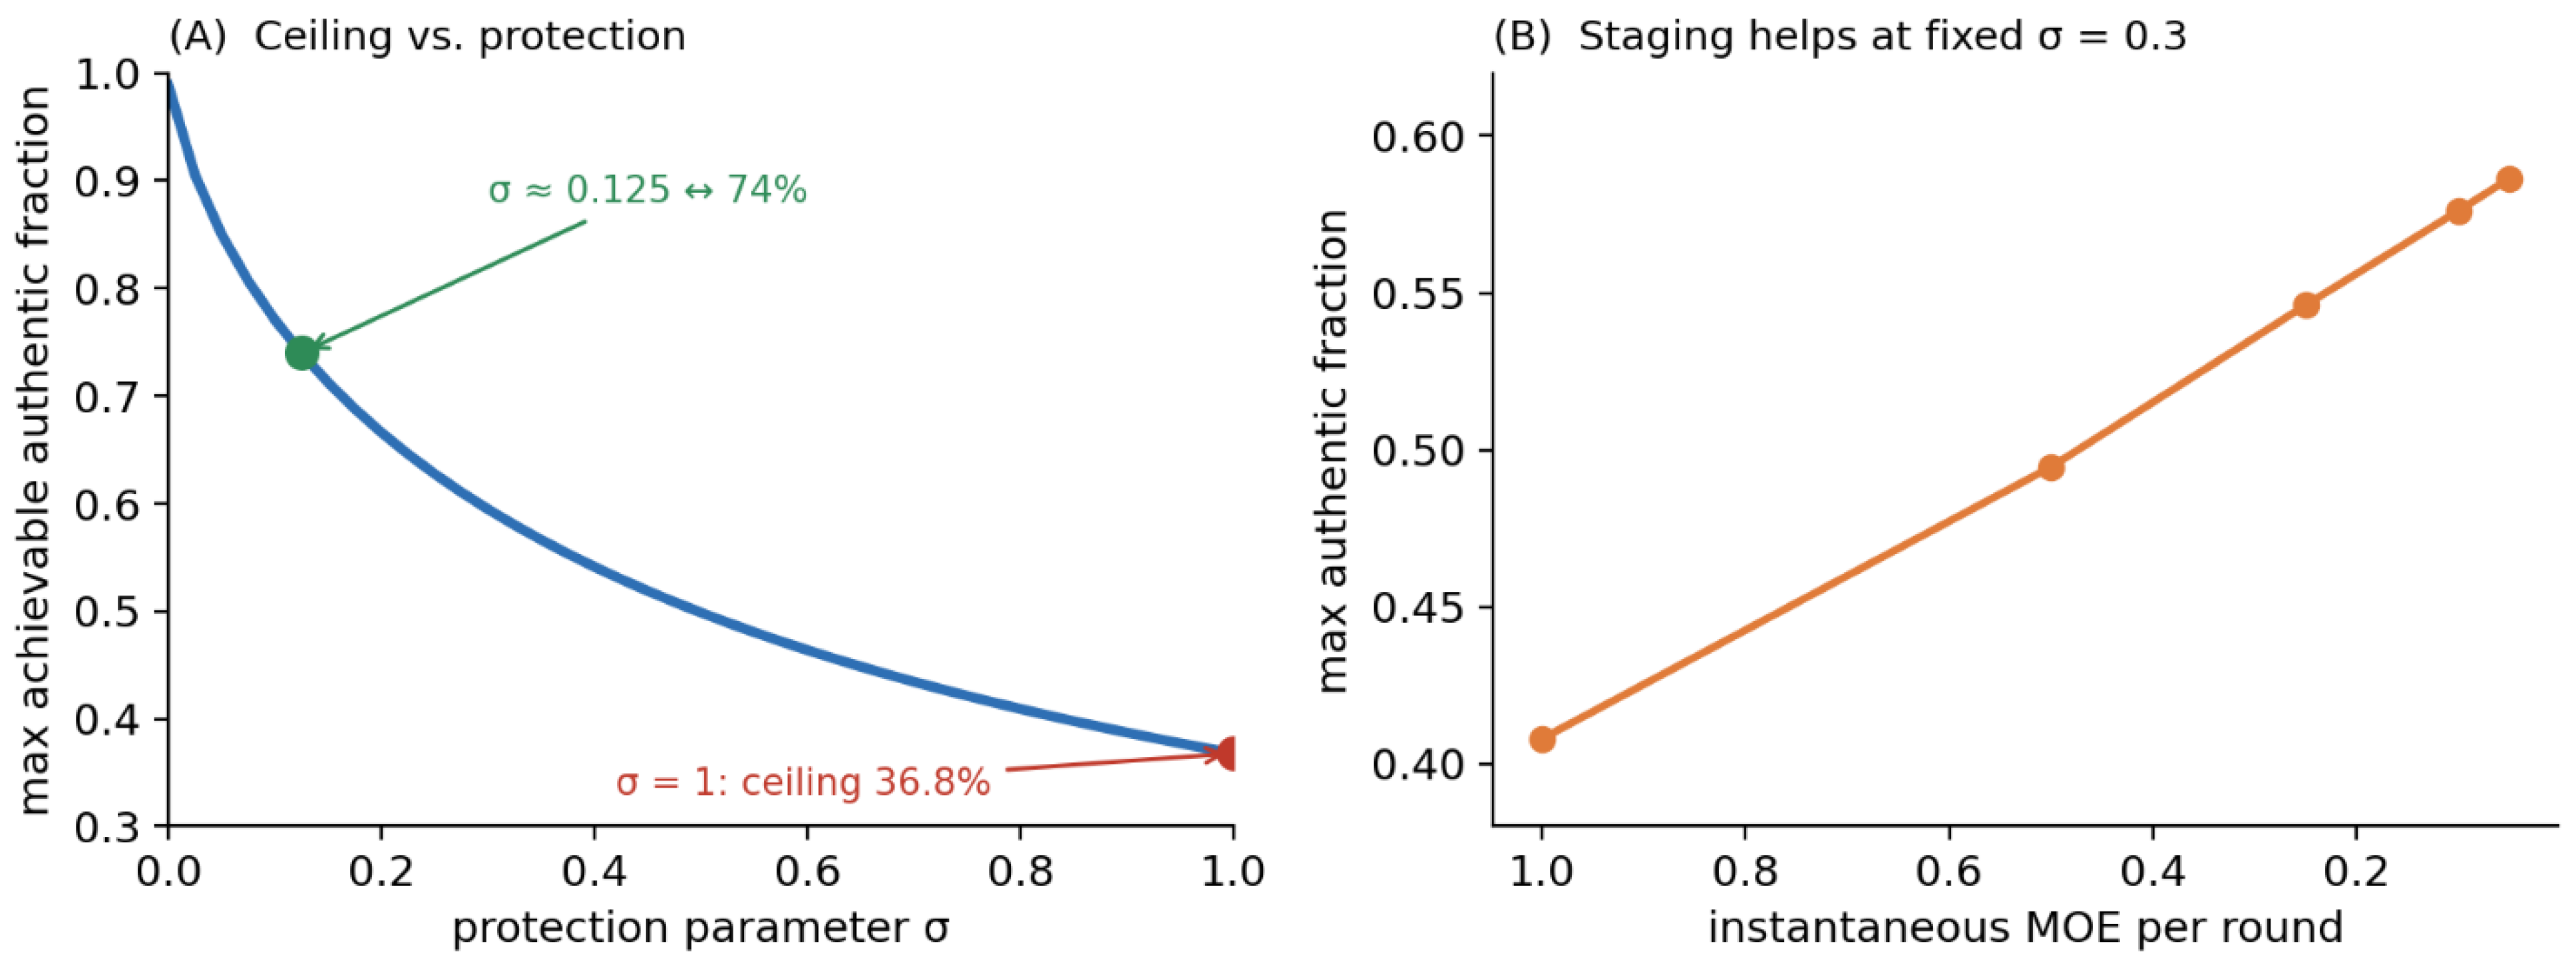
When σ = 1, staging the genome dose has no effect. A total MOE of 1 yields the same occupancy distribution whether delivered in one, ten, or one hundred rounds: 36.8% empty capsids, 36.8% authentic single-genome capsids, and 26.4% multi-occupancy particles. Across all genome-dosing schedules, the maximum authentic fraction remains exactly 1/e. Thus, when loading is memoryless (that is, when loading events are fully independent), only the total dose matters, and iterative genome addition (staging) cannot exceed the stochastic ceiling. For σ < 1, the achievable authentic fraction is a “smooth function” of σ: ~49.9% at σ = 0.5, ~59.5% at σ = 0.3, ~66.6% at σ = 0.2, ~77.1% at σ = 0.1, and approaching ~99% as σ → 0.
Staging improves authentic yield only when σ < 1, but under that condition the effect can be substantial. For example, at σ = 0.3, reducing the instantaneous MOE from 1.0 to 0.05 increases the achievable authentic fraction from about 41% to about 59%. This increase occurs because low per-round genome availability allows capsids to accumulate in the protected single-occupancy state faster than they transition to multi-occupancy.
Importantly, these values are not merely simulation output. In the fully staged limit (per-round dose approaching zero), the proposed model yields an exact closed-form solution: the maximum achievable authentic fraction equals [e^(−σλ*) − e^(−λ*)] / (1 − σ), achieved at the cumulative per-empty-capsid dose λ* = ln(1/σ)/(1 − σ). This expression reproduces the values above and recovers the 1/e ceiling as σ → 1. A full derivation is provided in Appendix C.
Figure 2.
(A) Maximum achievable authentic fraction as a function of the protection parameter σ; the σ = 1 limit yields the 36.8% ceiling, and σ ≈ 0.125 corresponds to ~74%. (B) At fixed partial protection (σ = 0.3), lowering the per-round instantaneous MOE increases the achievable authentic fraction.
This framework clarifies how a manufacturing strategy can reduce empty capsids without simply increasing defective full particles. The key intervention is not gradual genome addition alone but rather combining low-instantaneous-MOE loading with occupancy-dependent protection, so that capsids already carrying one genome are less likely to capture another. In the model, staged loading improves authentic yield only when σ < 1. Recent production studies are consistent with this type of control, although they do not measure σ directly.
For example, chronologically distributed transfection, in which capsid components and payload are delivered before the genome-replication machinery, increased capsid filling severalfold and supports the idea that capsids can assemble before genomes become abundant (Chen et al. 2025). Inducible producer systems likewise separate replication from packaging and allow tuning of the genome-containing fraction (Merten 2024, Ren et al. 2025). The occupancy model provides a quantitative interpretation: authentic yield should depend on σ, whereas interventions that increase only the total genome supply are predicted to raise the apparent full fraction mainly by increasing defective multi-occupancy particles.
As a calibration point, an authentic fraction near 74% corresponds to σ ≈ 0.125. In other words, once a capsid has captured one genome, it would need to be about 88% less likely than an empty capsid to capture another genome. This level of protection is physically plausible given the capsid’s approximately 1.25-genome-equivalent interior volume, but σ should be treated as an empirical parameter to be measured rather than as an assumed property of the system.
Appendix A presents a specific biological intervention strategy for implementing controlled, low-instantaneous-MOE loading.
4. Discussion
4.1. Multi-Channel Insertion: An Overview of the Mechanism
While the ceiling argument is mechanism-agnostic, it must still account for how defective particles might form and yield testable (falsifiable) predictions. I suggest that the primary pathway to stealth-defective multi-occupancy particles is the simultaneous or near-simultaneous insertion of DNA through more than one of the capsid’s twelve fivefold channels. Although prevailing single-portal models can explain individual translocation events, they often assume these events occur independently, ignoring the possibility that the other eleven channels could be involved. Currently, no strong evidence supports such an exclusion.
Several features relevant to AAV genome packaging are well established. Mature capsids have twelve channels at the fivefold vertices, and Rep-mediated genome translocation occurs through a fivefold pore; individual packaging events can also proceed processively from the 3′ end (Bleker et al. 2006, Bleker et al. 2005, Kaelber et al. 2025, King et al. 2001, Mietzsch et al. 2019). These observations support a single-portal pathway for an individual translocation event, but they do not, by themselves, show that a capsid engages only one pore or initiates packaging only once before sealing. Without a demonstrated exclusion mechanism, the presence of twelve equivalent, Rep-accessible channels leaves open the possibility that high genome availability could permit multi-channel engagement or repeated initiation on the same capsid. Consequently, the burden of proof falls on any claim that AAV packaging is obligatorily restricted to a single portal under all production conditions.
Two observations support this hypothesis. First, the single-particle data summarized in the Introduction demonstrate multi-equivalent and subgenomic occupancy; multi-occupancy occurs, leaving the mechanistic route as the remaining question. Second, the capsid interior can accommodate approximately 1.25 genome-equivalents. This capacity is sufficient to accommodate a plurality of short DNA species, each inserted through distinct channels, to contribute to a full-density particle without any one DNA species being an intact genome.
This hypothesis, while testable, is not required for defining the stochastic ceiling itself. If multi-channel insertion occurs, it yields three experimentally distinguishable predictions. First, capsids captured during active packaging, particularly under high genome availability, should sometimes show DNA or Rep motors associated with more than one fivefold channel. Second, the frequency of multi-occupancy signatures should increase with genome supply, as the Poisson model predicts. Third, AAV packaging should not require a strict structural or kinetic mechanism that limits each capsid to a single active portal. A direct demonstration of obligatory single-portal packaging would therefore falsify the multi-channel mechanism. It would not, however, eliminate the mechanism-agnostic ceiling described in Section 3.
Multi-occupancy could also arise through pathways other than simultaneous multi-channel insertion. These include sequential reloading before capsid sealing, co-packaging of snapback or replication intermediates, or partial loading followed by truncation. These mechanisms are compatible with the occupancy model and need not be excluded for the stochastic argument to hold. The model focuses on multi-channel insertion because the twelve fivefold channels provide a clear structural basis for the hypothesis and make it experimentally testable.
4.2. Aligning the Model with Reported Full-Capsid Fractions
A potential objection is that reported “percent full” values often exceed 36.8%. However, this objection conflates two distinct quantities and therefore does not invalidate the model.
First, the highest reported values, often 50–90% or higher, usually refer to material after density-gradient or ion-exchange enrichment, when empty capsids have been selectively depleted. The proposed production ceiling does not limit this kind of downstream enrichment; instead, it limits the fraction of the original capsid population expected to contain exactly one authentic genome. Thus, high “percent full” values after purification describe the composition of the enriched product, not the production-level yield of authentic single-genome capsids.
Second, and more importantly, “full” in a purified preparation often refers to capsids that are maximally filled, as indicated by density, charge, or another bulk physical property. As such, these measurements report total packaged material, not whether each capsid contains a single intact vector genome. A preparation described as 90% full could therefore include authentic single-genome capsids alongside stealth-defective particles containing partial, rearranged, or multiple DNA species. Such a result is consistent with an authentic fraction near the stochastic ceiling if the remaining full-density particles are defective.
Direct single-particle measurements offer a more relevant comparison because they count genome-containing capsids before enrichment, rather than inferring fullness from net bulk properties. In unpurified preparations using a standard Rep system, cryo-EM studies reported full-capsid fractions of approximately 10–20%; engineered Rep improvements increased this value only to the low-to-mid 30% range (Mietzsch et al. 2021). These production-level values remain below 36.8%, consistent with the proposed ceiling and the view that direct single-particle counts cluster beneath the predicted bound.
4.3. The Measurement Problem and a Conclusive Test
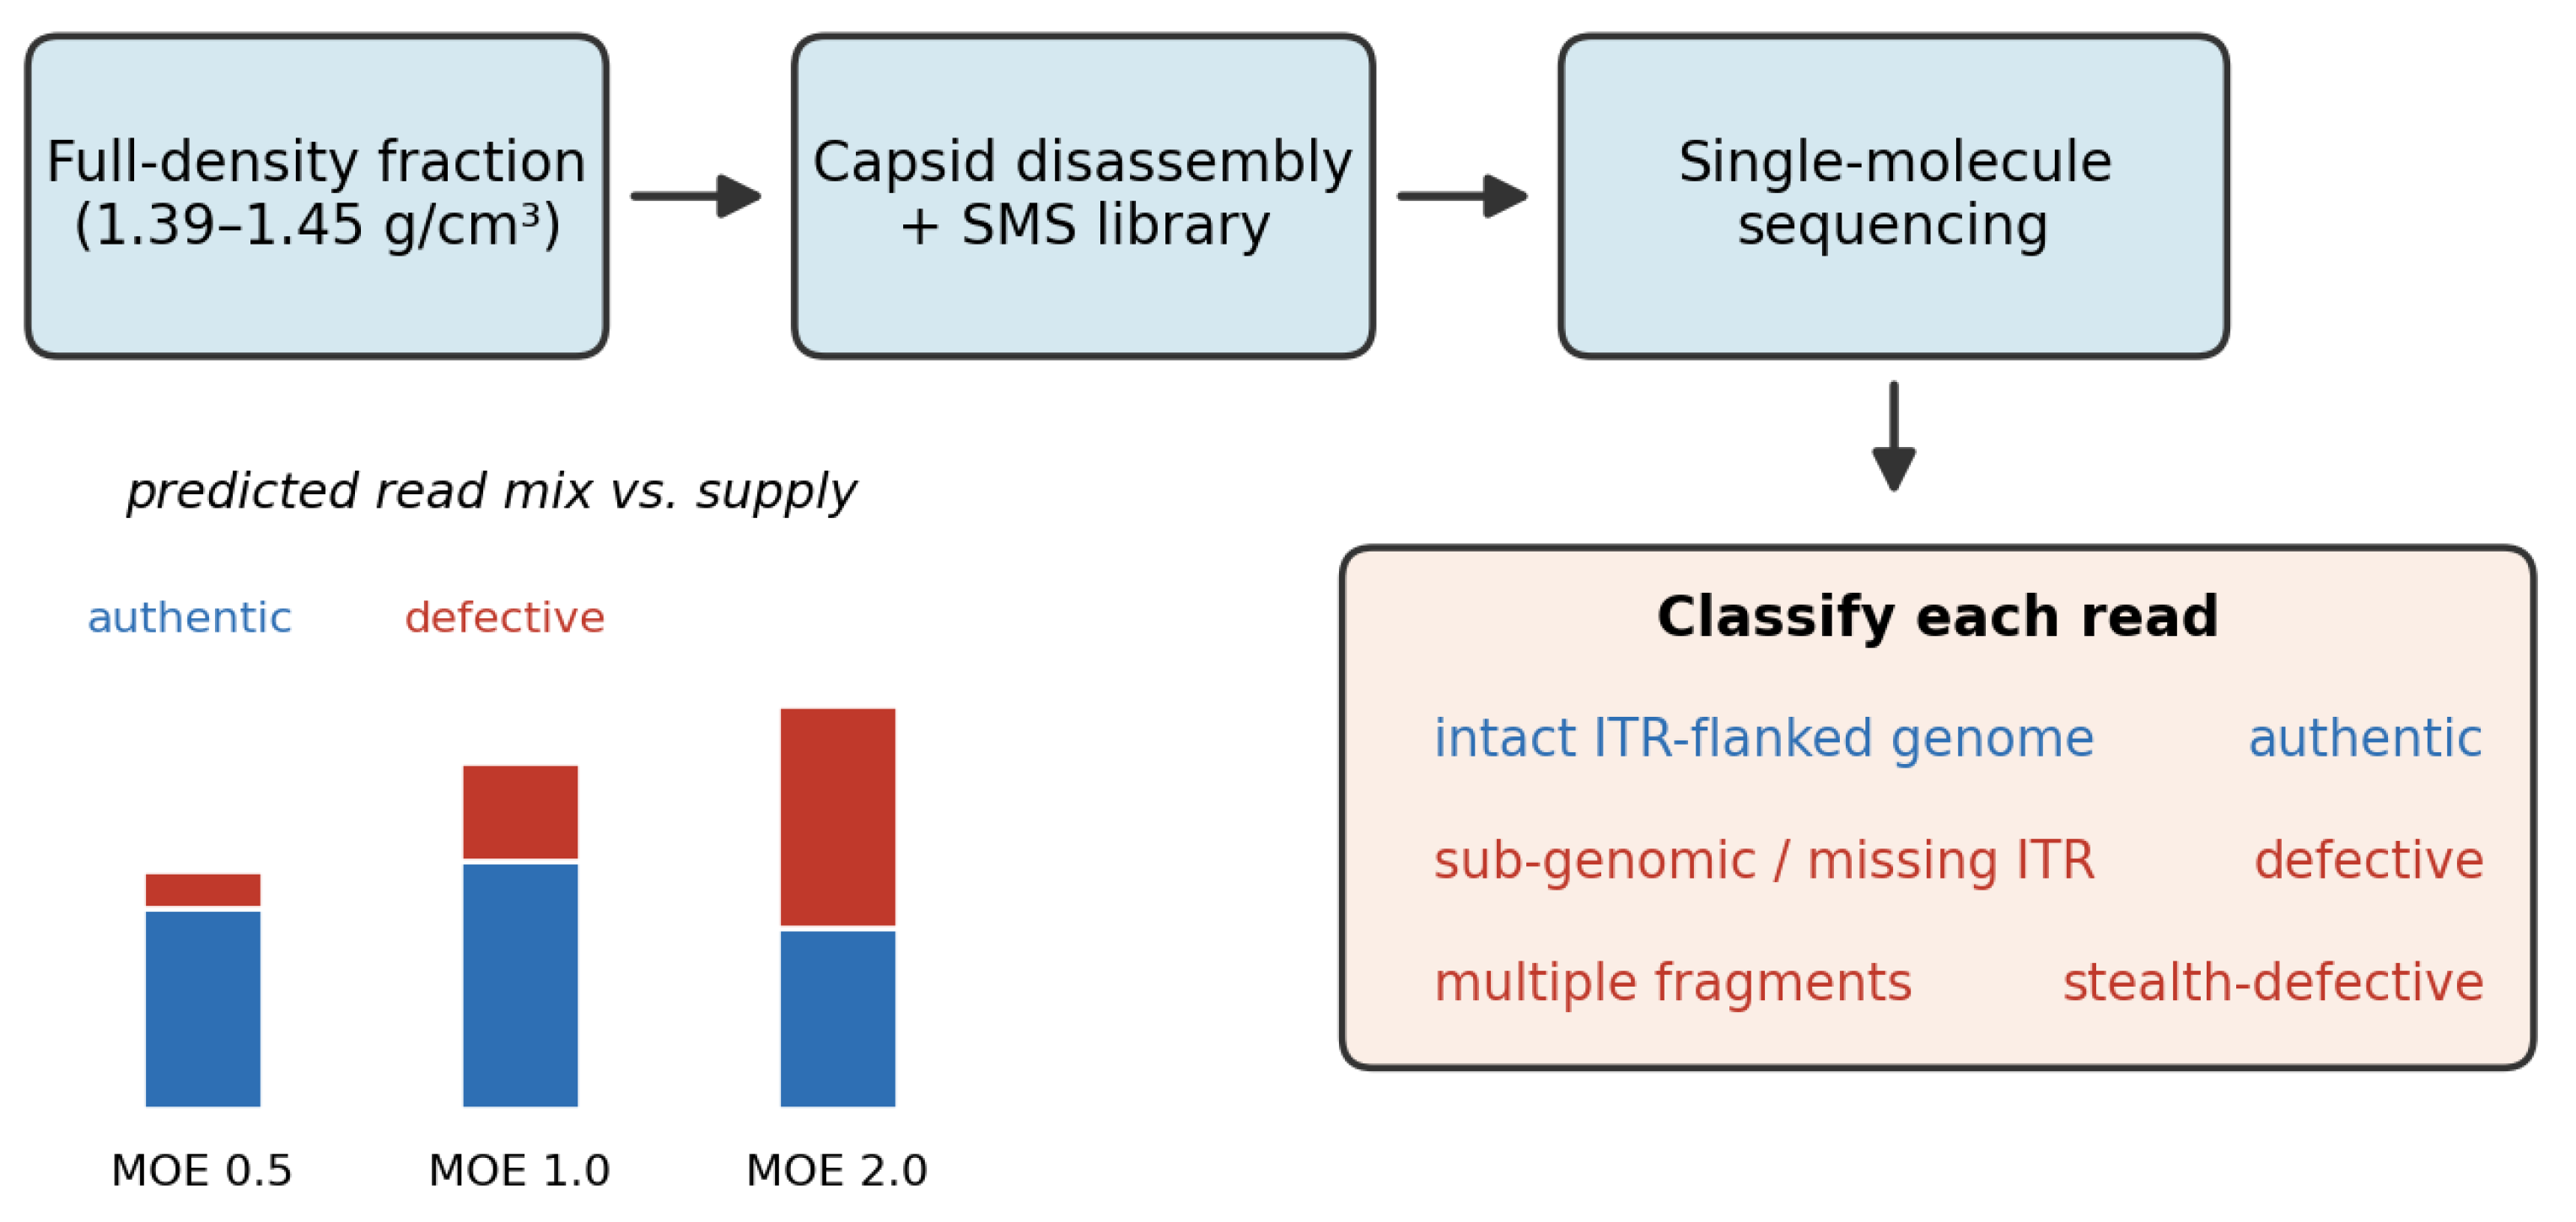
Conventional assays such as density measurement, light scattering, electron microscopy, and production-batch genome titering can identify particles containing packaged DNA, but they cannot determine whether that DNA is a single intact vector genome or an equivalent mass of partial fragments. As a result, these methods cannot quantify the authentic fraction. The most direct approach is single-molecule sequencing of the full-density fraction, which can classify molecules as intact, ITR-flanked, full-length genomes; subgenomic fragments lacking one or both ITRs; or sequence patterns consistent with multi-occupancy. The Kapranov (Kapranov et al. 2012) study summarized above provides precedent for applying this strategy to purified full-density rAAV particles.
The decisive experiment is to sequence the full-density fraction across a controlled genome-to-capsid series and compare the resulting authentic and subgenomic fractions with the occupancy model. Under independent batch loading, the model predicts that authentic single-genome capsids should peak near MOE = 1 and decline as genome supply increases, while subgenomic and multi-occupancy signatures should become more abundant. It also predicts that the authentic fraction should not exceed approximately 36.8% in this regime. In contrast, if the authentic fraction within the full-density band rises continuously toward 100% with increasing genome supply, the framework would be falsified.
Figure 3.
Single-molecule sequencing assay to discriminate authentic from stealth-defective particles, with predicted read-class distributions at MOE values of 0.5, 1.0, and 2.0.
Figure 3.
Single-molecule sequencing assay to discriminate authentic from stealth-defective particles, with predicted read-class distributions at MOE values of 0.5, 1.0, and 2.0.
4.4. Implications
If this framework is correct, reported authentic full-capsid fractions far above approximately 37% in single-batch production should be interpreted as mixtures of authentic and stealth-defective particles in unknown proportions. Strategies that increase total genome supply while keeping capsid expression fixed are predicted to enrich defective particles rather than authentic vectors. Potency and dose calculations based on total vector genomes may therefore overestimate the transduction-competent dose to the extent that stealth-defective particles contribute genome-equivalents. Conventional lot-release assays cannot confirm the authentic fraction, defined here as capsids containing exactly one intact genome. These implications do not depend on the specific multi-channel mechanism proposed in Section 4.1; they follow from stochastic multi-occupancy and bulk methods' inability to distinguish authentic from stealth-defective particles. The multi-channel hypothesis is offered as the most direct and experimentally testable explanation for the emergence of multi-occupancy.
4.5. Discussion Summary and Future Model Validation
This framework reframes an established quality metric as the observable outcome of a constrained stochastic process. Its strength lies in combining elementary occupancy mathematics with single-particle evidence already accepted in the field. The central empirical claim—that the full-density fraction is heterogeneous and that the authentic fraction within it is bounded near 1/e under independent batch loading—is directly testable and already partially supported. The extent to which the bound can be exceeded reduces to a single measurable parameter, σ.
Several parts of the framework remain untested. Section 4.1 proposes a multi-channel mechanism intended to be both impactful and easily falsified if incorrect: it challenges the assumption that AAV packaging is necessarily limited to one active portal per capsid. However, the stochastic ceiling derived in Section 3 does not depend on that mechanism and would still apply even if multi-channel insertion were disproven. Likewise, the Poisson model treats loading events as independent, an idealization that can be relaxed by measuring occupancy-dependent protection through σ. The proposed single-molecule sequencing experiment is designed to resolve the remaining empirical question: how packaged fragments, partial genomes, and genome-equivalent occupancy relate to authentic single-genome capsids. Thus, the 36.8% value should be understood as the null expectation for independent stochastic loading, not as a measured constant.
This framework complements, rather than duplicates, adjacent work. Native mass spectrometry has shown that AAV capsid assembly is stochastic with respect to VP composition (Wörner et al. 2021). The present model addresses a different stochastic step: genome loading into assembled capsids. Similarly, prior single-molecule and charge-detection studies have established that packaged genomes are heterogeneous. This work links that heterogeneity to a quantitative occupancy bound, defines σ as a measurable means to exceed the bound, proposes multi-channel insertion as a testable mechanism, and identifies single-molecule sequencing of the full-density fraction as the decisive experimental test.
5. Conclusions
This work proposes that authentic full-capsid yield in single-batch rAAV production is bounded near 36.8% by stochastic genome loading. Increasing genome supply beyond the optimum is predicted to convert empty capsids into physically full but functionally defective multi-occupancy particles that conventional analytics cannot resolve. Direct single-particle counts are consistent with this bound, whereas higher reported full-capsid values often reflect post-purification enrichment and cannot establish the authentic fraction without sequence-level analysis. I further propose that multi-channel insertion is a principal mechanism for generating these defective particles and that single-portal exclusivity remains an unproven assumption. Single-molecule sequencing of the full-density fraction across varied genome supplies provides the decisive test of the framework. Within this model, authentic yield is improved not by maximizing genome supply, but by exploiting occupancy-dependent protection, represented by σ, through decoupled, low-instantaneous-MOE loading. The framework therefore challenges the interpretation of a widely used quality metric while identifying a measurable ceiling, a testable mechanism, and a practical route for improving authentic vector yield.
Appendix A. A Potential Biological Throttle: The FKBP52 / ITR D-Sequence Interaction
Section 3.4 showed that exceeding the stochastic ceiling requires two linked conditions: a loaded capsid must be less likely than an empty one to capture another genome (σ < 1), and packageable genomes must be supplied slowly enough for capsids to accumulate in the protected single-occupancy state. Physical approaches, such as staged delivery of replication components or inducible control of genome replication, provide direct ways to lower instantaneous genome availability. Here I consider a more speculative alternative: a pharmacologically tunable biological control that could continuously meter genome availability, acting as a rheostat rather than an on/off switch. The individual components of this proposal are established in the AAV literature; what remains unproven is whether they can be integrated to control genome supply during vector production.
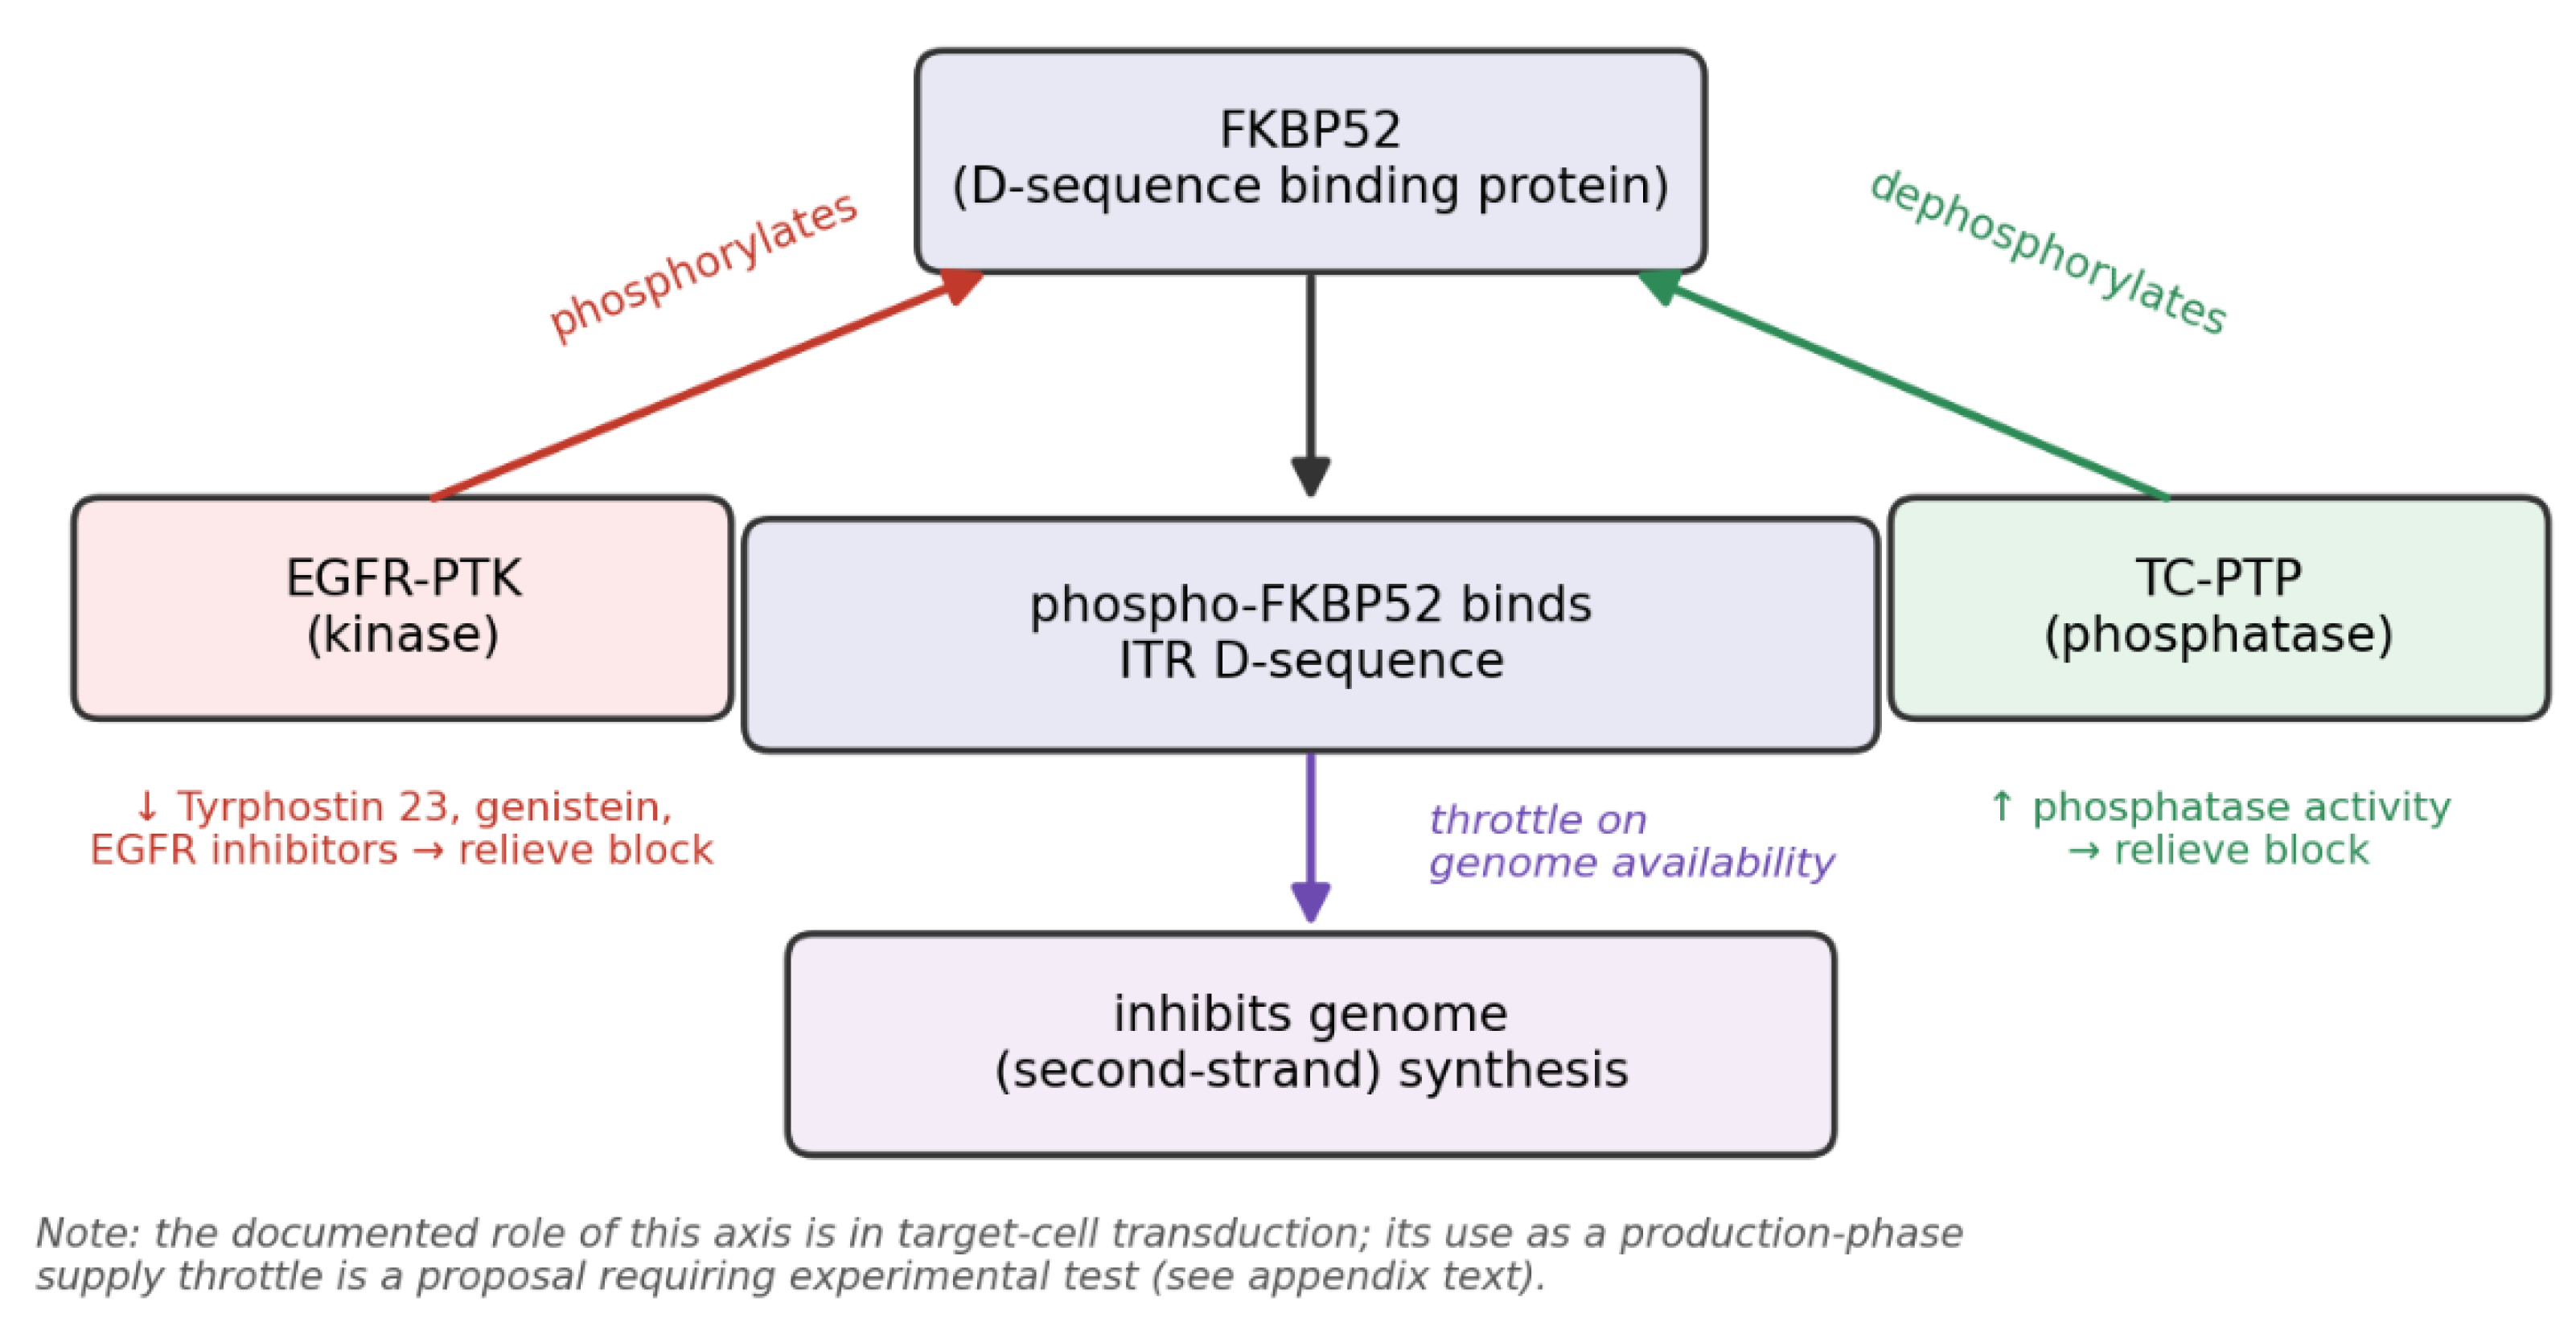
The candidate control point is the interaction between FKBP52 and the AAV inverted terminal repeat (ITR). Each ITR contains a short single-stranded D-sequence that is involved in genome replication and packaging. FKBP52 binds this D-sequence in a phosphorylation-dependent manner: the tyrosine-phosphorylated form binds the D-sequence and inhibits conversion of the single-stranded vector genome to double-stranded DNA during second-strand synthesis, whereas the dephosphorylated form does not (Mah et al. 1998, Qing et al. 2001). The D-sequence is therefore both a structural element of the AAV genome and a regulatable site in AAV DNA metabolism.
Opposing enzymes regulate this interaction. The EGFR protein tyrosine kinase phosphorylates FKBP52, favoring the inhibitory state, whereas T-cell protein tyrosine phosphatase removes the phosphate, relieving the block (Qing et al. 2003, Zhao et al. 2006). Because small molecules, including Tyrphostin 23, genistein, and clinically used EGFR inhibitors, can shift this phosphorylation state, the pathway is pharmacologically accessible (Zhong et al. 2007). In principle, such dose-dependent control could provide an external handle on a D-sequence-dependent step.
The proposed application is to use this type of control to regulate the rate at which packageable genomes accumulate in producer cells without directly altering capsid expression. If genome availability could be slowed and titrated in this way, the production system could approximate the low-instantaneous-MOE regime described in Section 3.4. Capsids would encounter genomes gradually, allowing protected single-occupancy particles to accumulate, and the achievable authentic fraction would then depend on σ rather than on bulk genome excess that drives multi-occupancy. A continuously tunable small-molecule rheostat would be operationally attractive because it could decouple genome supply from capsid supply without requiring repeated transfection or a dedicated inducible production line.
This proposal remains speculative. The FKBP52 studies cited above were conducted in target-cell transduction, where FKBP52 influences second-strand synthesis after vector entry. They did not test producer cells, where the relevant variables are Rep-mediated replication and the accumulation of single-stranded genomes available for packaging. Using the FKBP52/D-sequence axis as a production-phase throttle therefore rests on an untested premise: that this interaction, or an engineered analog, can modulate the supply of packageable genomes during manufacturing. The first proof-of-concept experiment would be to determine whether pharmacological modulation of FKBP52 phosphorylation alters the rate at which packageable genomes accumulate in producer cells.
Several practical questions would also need to be resolved. The modulation would need to slow, rather than accelerate, genome availability; the required small-molecule concentrations would need to preserve producer-cell viability and overall yield; and the timing of treatment would need to be coordinated with capsid assembly. For these reasons, this approach is less immediately actionable than staged delivery or inducible replication, which more directly decouple genome production from capsid production. It is included because, if the premise is validated, it could provide a simple, continuously tunable implementation of the loading control required by the model using experimentally accessible molecules.
Figure A1.
The FKBP52 / ITR D-sequence axis as a candidate genome-supply throttle. The epidermal growth factor receptor protein tyrosine kinase (EGFR-PTK) phosphorylates FKBP52; the phosphorylated form binds the single-stranded D-sequence of the AAV ITR and inhibits genome (second-strand) synthesis, whereas T-cell protein tyrosine phosphatase (TC-PTP) reverses this modification and relieves the block. Small-molecule EGFR-PTK inhibitors (e.g., Tyrphostin 23, genistein) shift the balance toward the non-inhibitory state, providing dose-dependent external control of the D-sequence-dependent step. This axis is documented to regulate target-cell transduction; I propose its use as a production-phase throttle on the supply of packageable genome, though this remains to be tested.
Appendix B. The Stochastic-Occupancy Ceiling,
Premises.
- Genome loading deposits genome equivalents into a population of preformed capsids.
- Loading events are independent and share a common mean rate; let
denote the mean number of genome-equivalents loaded per capsid (the multiplicity of encapsidation, or MOE).
- 3.
- No capsid-level memory: the probability that a capsid receives its next genome-equivalent is independent of how many it already holds. (This premise is relaxed in the Appendix C. Premises.)
Derivation. Premises 1–3 define a Poisson process, so the number of genome-equivalents in a given capsid is Poisson-distributed.
The three operationally relevant occupancy classes are:
The multi-occupancy term is simply the tail of the series, .
The authentic single-genome fraction is . Differentiating,
and the second derivative at confirms a maximum. Therefore,
At this optimum, the population is empty and multi-occupancy.
Monotonic decline beyond the optimum. For , , so raising genome supply beyond one equivalent per capsid strictly lowers the authentic fraction: the added genomes convert empty capsids into multi-occupancy particles rather than authentic ones. As a polynomial expansion,
a series in whose coefficients are the alternating reciprocal factorials.
Appendix C. Occupancy-Dependent Protection: The Model and the Achievable Ceiling
This appendix relaxes the constraint in premise 3 of Appendix B to specify how far, and under what conditions, the ceiling can be exceeded.
Premises.
- Empty capsids capture genome equivalents via independent Poisson loading, as in Appendix B.
- capsid that already holds one genome captures a second genome at a rate times that of an empty capsid, with . Here, indicates no protection (full memorylessness), and indicates complete protection. As introduced here, is the relative second-capture rate; it is a defined protection coefficient, not a quantity derived from the Poisson model.
- Loading is delivered over rounds; in each round, an empty capsid receives a mean dose “” (the instantaneous MOE per round), and a single-occupancy capsid receives a mean dose .
Discrete recursion. Writing for the empty, single, and multi fractions after each round and applying the within-round Poisson splits (an empty capsid stays empty with probability , gains exactly one with , and gains two or more with ; a single-occupancy capsid stays single with and leaks to multi with ),
The invariance (staging does nothing). When the single-occupancy capture rate equals the empty-capsid rate, the population is indistinguishable from one-shot Poisson loading with total mean . Then , maximized at to give exactly , for any schedule. Iterative or staged addition therefore cannot beat the ceiling unless .
Continuous (fully-staged) limit and closed-form model. Let the per-round dose with the cumulative empty-dose retained as the loading variable. The probability of gaining two or more genomes in an infinitesimal increment is , so empties flow only to singles, and the recursion becomes a linear system.
Hence and, solving the first-order linear equation for ,
The limit recovers (by L’Hôpital rule), consistent with Appendix B.
Maximization. Setting ,
and the achievable authentic-fraction ceiling as a function of protection is
Numerical values (by direct substitution; the discrete recursion with fine staging agrees to better than ):
| 1.000 | 1.000 | 0.368 |
| 0.500 | 1.386 | 0.500 |
| 0.300 | 1.720 | 0.597 |
| 0.200 | 2.012 | 0.669 |
| 0.125 | 2.377 | 0.743 |
| 0.100 | 2.558 | 0.774 |
Thus, the figure corresponds to : a loaded capsid roughly less likely to capture a second genome than an empty one.
Polynomial-expansion form. Expanding the closed form about ,
a power series in whose coefficients are the finite geometric sums .
At , each such sum equals , reproducing ; note, these are the explicit σ-polynomial coefficients derived through automated symbolic computation software.
Author Contributions
G.L.D.: Conceptualization, theory development, mathematical analysis, manuscript writing.
Conflicts of interest
The author declares no conflicts of interest.
Declaration of generative AI and AI-assisted technologies in manuscript preparation
During preparation of this manuscript, the author used Claude Opus 4.8 (Anthropic) to convert hand-derived mathematical expressions into compact symbolic notation, assist in extending and solving the model under the author’s direction, and generate computational figures. After using this tool, the author reviewed and edited all AI-assisted content as needed and takes full responsibility for the final manuscript.
Funding
This research received no external funding.
Competing interests
The author declares no competing interests.
Data availability statement
This is a theoretical manuscript. No new experimental data were generated or analyzed. All mathematical derivations underlying the results are presented in full in Appendices B and C. Additional intermediate derivation steps, symbolic-computation output, and the parameter-space calculations underlying Figures 1 and 2 are available from the author upon reasonable request.
References
- Bleker, S.; Pawlita, M.; Kleinschmidt, J.A. Impact of capsid conformation and Rep-capsid interactions on adeno-associated virus type 2 genome packaging. J. Virol. 2006, 80, 810–20. [Google Scholar] [CrossRef]
- Bleker, S.; Sonntag, F.; Kleinschmidt, J.A. Mutational analysis of narrow pores at the fivefold symmetry axes of adeno-associated virus type 2 capsids reveals a dual role in genome packaging and activation of phospholipase A2 activity. J. Virol. 2005, 79, 2528–40. [Google Scholar] [CrossRef]
- Chen, Q.; Lee, C.H.; Whitfield, R.; Nesbeth, D.N. Chronologically distributed transfection improves AAV2 and AAV2/8 capsid filling and reveals assembly schedule divergence. Mol. Ther. Methods Clin. Dev. 2025, 33, 101610. [Google Scholar] [CrossRef]
- Kaelber, J.T.; Barnakov, V.; Shen, J.; Hernandez, K.; Tarbox, H.J.; Khan, A.; Escalante, C.R. Insights into the AAV packaging mechanism: Cryo-EM Structure of the AAV2 Rep-Capsid Packaging Complex. bioRxiv 2025. [Google Scholar] [CrossRef]
- Kapranov, P.; Chen, L.; Dederich, D.; Dong, B.; He, J.; Steinmann, K.E. Xiao W (2012) Native molecular state of adeno-associated viral vectors revealed by single-molecule sequencing. Hum. Gene Ther. 23, 46–55. [CrossRef]
- King, J.A.; Dubielzig, R.; Grimm, D.; Kleinschmidt, J.A. DNA helicase-mediated packaging of adeno-associated virus type 2 genomes into preformed capsids. EMBO J. 2001, 20, 3282–91. [Google Scholar] [CrossRef]
- Kuzmin, D.A.; Shutova, M.V.; Johnston, N.R.; Smith, O.P.; Fedorin, V.V.; Kukushkin, Y.S. Johnstone EC (2021) The clinical landscape for AAV gene therapies. Nat. Rev. Drug Discov. 20, 173–174. [CrossRef]
- Lecomte, E.; Tournaire, B.; Cogne, B.; Dupont, J.B.; Lindenbaum, P.; Martin-Fontaine, M. Leger A (2015) Advanced Characterization of DNA Molecules in rAAV Vector Preparations by Single-stranded Virus Next-generation Sequencing. Mol. Ther. Nucleic Acids 4, e260. [CrossRef]
- Mah, C.; Qing, K.; Khuntirat, B.; Ponnazhagan, S.; Wang, X.S.; Kube, D.M. Srivastava A (1998) Adeno-associated virus type 2-mediated gene transfer: role of epidermal growth factor receptor protein tyrosine kinase in transgene expression. J. Virol. 72, 9835–43. [CrossRef]
- Merten, O.W. Development of Stable Packaging and Producer Cell Lines for the Production of AAV Vectors. Microorganisms 2024, 12. [Google Scholar] [CrossRef]
- Mietzsch, M.; Eddington, C.; Jose, A.; Hsi, J.; Chipman, P.; Henley, T. Agbandje-McKenna M (2021) Improved Genome Packaging Efficiency of Adeno-associated Virus Vectors Using Rep Hybrids. J. Virol. 95, e0077321. [CrossRef]
- Mietzsch, M.; Penzes, J.J.; Agbandje-McKenna, M. Twenty-Five Years of Structural Parvovirology. Viruses 2019, 11. [Google Scholar] [CrossRef]
- Nakatsuka, R.; Yamaguchi, Y.; Hirohata, K.; Shimojo, S.; Murakami, M.; Rocafort, M.A.V. Uchiyama S Multimass Analysis of Adeno-Associated Virus Vectors by Orbitrap-Based Charge Detection Mass Spectrometry. Anal. Chem. 2024, 96, 17037–17046. [Google Scholar] [CrossRef]
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2026 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (http://creativecommons.org/licenses/by/4.0/).
Copyright: This open access article is published under a Creative Commons CC BY 4.0 license, which permit the free download, distribution, and reuse, provided that the author and preprint are cited in any reuse.