

Submitted:
25 December 2025
Posted:
26 December 2025
You are already at the latest version
Abstract
Background: Schizophrenia is a highly heritable disorder with a polygenic architecture, and growing evidence points to glutamatergic dysfunction—particularly involving NMDA receptor hypofunction and impaired synaptic plasticity—as a key mechanistic contributor. The latest Psychiatric Genomics Consortium (PGC3) genome-wide association study reinforced synaptic biology as a major locus of common variant risk.Methods: Using MAGMA gene-set analysis, we examined polygenic enrichment in the PGC3 European-ancestry schizophrenia GWAS summary statistics (53,386 cases, 77,258 controls). Two hypothesis-driven glutamatergic/synaptic plasticity gene sets were tested: a narrow core set (23 genes) focused on ionotropic receptors and direct modulators, and an expanded set (130 genes) that additionally included transporters, metabolic enzymes, scaffolding proteins, and downstream plasticity cascades (BDNF-TrkB, mTOR, CREB, immediate-early genes). Monoaminergic (104 genes) and housekeeping (184 genes) sets served as negative controls. Competitive gene-set testing was performed with Bonferroni correction and false-discovery rate estimation.Results: The expanded glutamate/plasticity set showed significant enrichment for schizophrenia polygenic signal (p = 0.0033; Bonferroni-corrected p = 0.0134; FDR ≈ 0.013), with a higher mean association strength than the genome-wide average. The narrower core set displayed only a trend (p = 0.092). Neither control set was enriched (monoaminergic p = 0.186; housekeeping p = 0.152). Post-hoc exploration highlighted contributions from intracellular regulators, including mTOR pathway components, CREB targets, and immediate-early genes.Conclusions: Common risk variants in schizophrenia converge preferentially on broad glutamatergic signalling and downstream synaptic plasticity mechanisms rather than solely on surface receptors or monoaminergic pathways. These findings support the development of therapeutic strategies targeting synaptic plasticity to address persistent cognitive deficits in the disorder.
Keywords:
schizophrenia
; cognition
; NMDA
; AMPA
; plasticity
Introduction
Schizophrenia is a chronic, highly disabling psychiatric disorder that affects roughly one in every hundred people worldwide. Family and twin studies place the heritability of the illness between 70% and 80% [1], and successive genome-wide association studies (GWAS) have confirmed a markedly polygenic architecture, with hundreds of common variants contributing small effects [2].
One influential mechanistic theory proposes that reduced activity of the N-methyl-D-aspartate (NMDA) receptor underlies many aspects of the disorder. Sub-anaesthetic doses of ketamine, a non-competitive NMDA antagonist, reproduce transient psychotic and cognitive symptoms in healthy volunteers [3]. Post-mortem work has documented diminished NMDA receptor scaffolding proteins such as PSD-95 and SAP102 and disrupted AMPA receptor trafficking in cortical tissue from people with schizophrenia [4,5].
The most recent GWAS from the Psychiatric Genomics Consortium (PGC3) supported this point of view. The NMDA subunit gene GRIN2A was found to be a high-priority locus in synaptic biology, showing convergence with rare disruptive variants [2]. One of the least responsive symptom domains is cognitive deficits, which are often present before the illness starts and last through different stages [6]. Bipolar disorder also has similar but less severe deficits, which raises questions about mechanisms that are common to both disorders and those that are unique to each [7].
Recent anecdotal evidence has helped revive attention to glutamatergic strategies for cognitive remediation in psychotic disorders. Cheung [8] detailed a single-case intervention in a young man with schizoaffective disorder who received add-on treatment with low-dose dextromethorphan—an NMDA-receptor antagonist—together with piracetam, an agent that enhances AMPA-mediated transmission, and Deanxit, which supplies both mild noradrenergic activity and CYP2D6 inhibition. Within a short period the patient reported noticeably “clearer” thought processes, improved attentional capacity, and better academic functioning, and these gains occurred without any worsening of mood or psychotic symptoms.
Against this background, the present study used MAGMA gene-set analysis [9] to examine polygenic enrichment within predefined glutamatergic and synaptic plasticity pathways in the PGC3 GWAS. Two glutamate-focused sets—one narrowly centred on ionotropic receptors and one broader set that also included transporters, metabolic enzymes, and downstream plasticity cascades (BDNF-TrkB, mTOR, CREB)—were tested. Monoaminergic genes and generic housekeeping genes served as negative controls to evaluate specificity.
Methods
Participants and GWAS summary statistics
We analysed the European-ancestry summary statistics released by the Psychiatric Genomics Consortium for schizophrenia (PGC Wave 3, autosomes only). After quality control the dataset comprised 53,386 cases and 77,258 controls, giving an effective sample size of ≈ 58,749 [2]. Only SNPs present in the European panel of the 1000 Genomes Project were retained for downstream work.
Gene-Based Association Testing
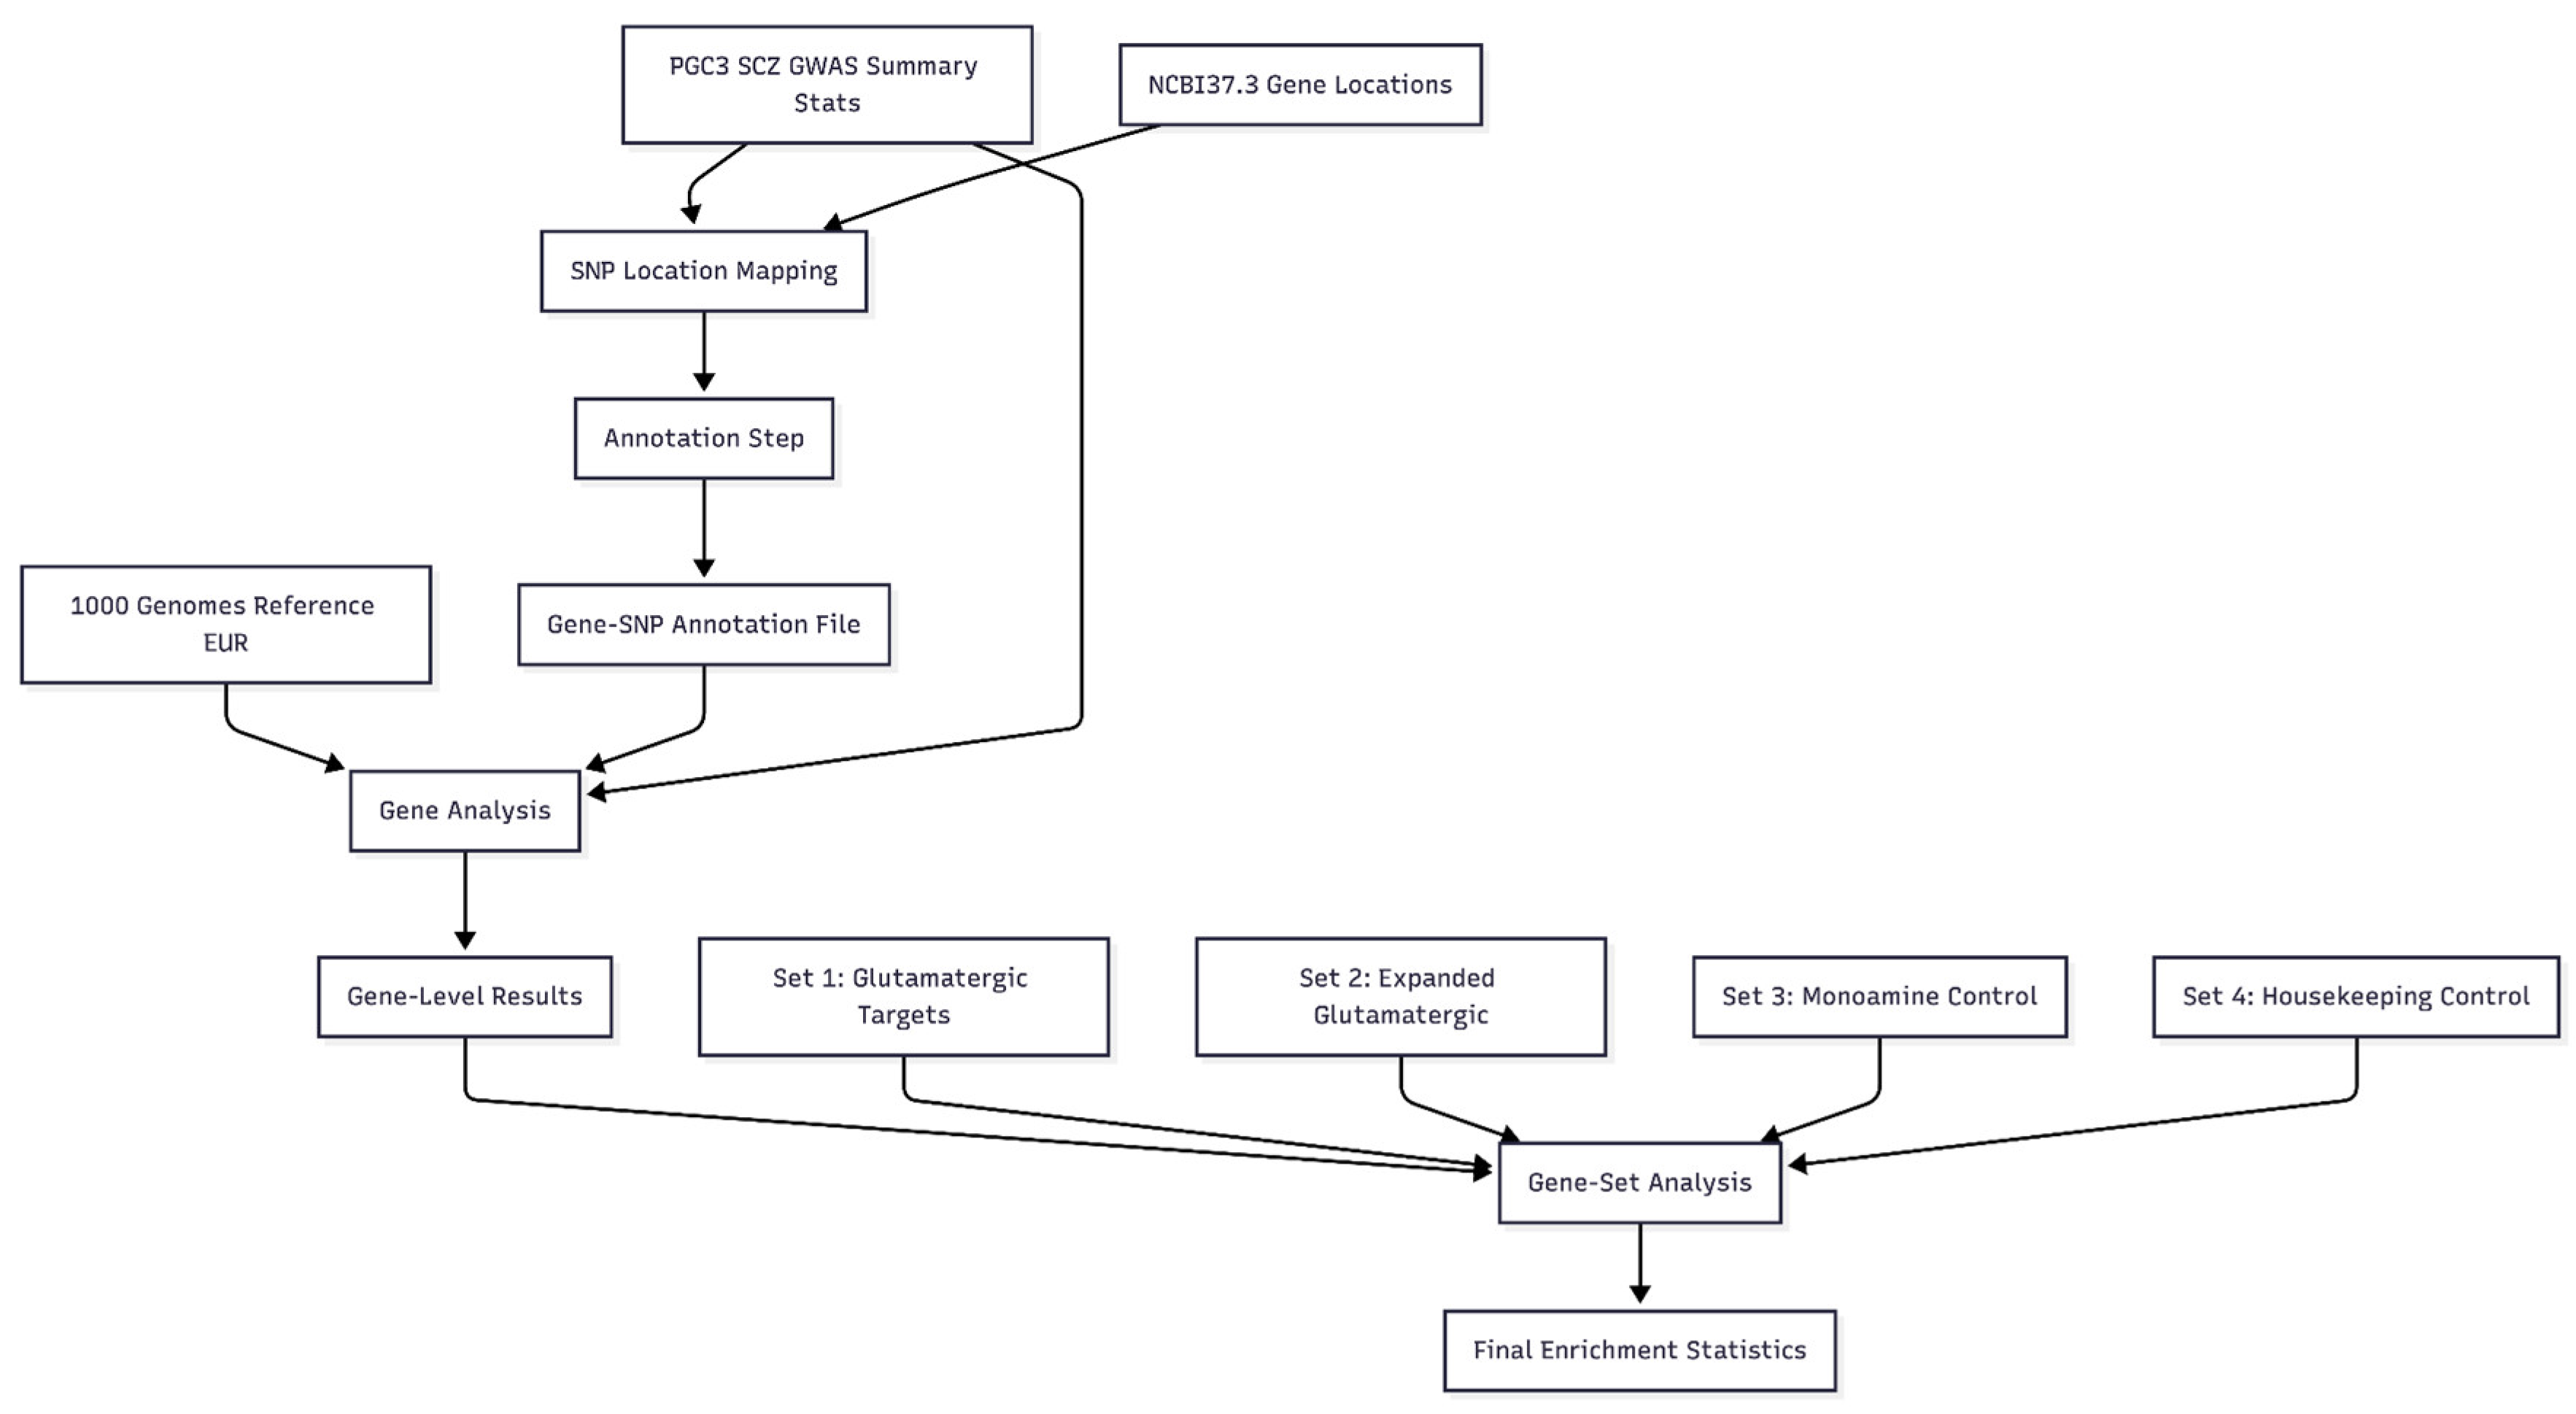
Gene-wise statistics were computed with MAGMA v1.10 [9]. Single-nucleotide variants were assigned to genes if they lay within 35 kb upstream or 10 kb downstream of the transcript boundaries defined in NCBI build 37 (hg19). Linkage disequilibrium was modelled with the European 1000 Genomes reference (phase 3). The SNP-wise mean model, with the PGC effective N as input, produced p-values for 18,449 genes.
Figure 1.
MAGMA Analysis Pipeline. The flowchart below illustrates the data processing steps applied to the PGC3 Schizophrenia GWAS summary statistics.
Figure 1.
MAGMA Analysis Pipeline. The flowchart below illustrates the data processing steps applied to the PGC3 Schizophrenia GWAS summary statistics.
Gene-Set Definitions
Four prespecified gene sets were evaluated.
- Candidate glutamate/plasticity core (23 genes) correspond to the targets of NMDA-AMPA modulation therapies, plasticity mediators and metabolic enzymes.
- Expanded glutamate/plasticity (130 genes) added all ionotropic receptor subunits, associated scaffolding proteins, transporters, metabolic enzymes, BDNF-TrkB, full mTOR and CREB cascades, immediate-early genes, CaMKII, PKA/PKC components and additional sigma receptors.
- Monoamine control (104 genes) covered receptors, transporters, and metabolic enzymes for dopamine, serotonin, noradrenaline, histamine and trace amines.
- House-keeping control (184 genes) comprised broadly expressed ribosomal, cytoskeletal, glycolytic, mitochondrial and heat-shock proteins.
Official gene symbols were converted to Entrez identifiers with the NCBI37.3 annotation file. After mapping, 22, 127, 96 and 160 genes, respectively, remained in the analysis.
Statistical Analysis
Genome-wide significance for individual genes was set at p < 2.71 × 10⁻⁶ (0.05/18,449). For sets, we compared the mean Z-score of member genes with the genome-wide mean using one-sample, one-sided t tests. Four sets were tested, so the Bonferroni threshold was p < 0.0125; false-discovery rates (FDR) were also calculated. Where appropriate, Kruskal–Wallis and Mann–Whitney procedures contrasted Z-score distributions. Analyses were run in Python 3 (pandas/numpy/scipy).
Results
Genome-Wide Gene Associations
The MAGMA run identified 705 genes exceeding the Bonferroni-corrected threshold and 6,059 additional genes at nominal p < 0.05. Top signals clustered in the extended MHC (e.g., BTNL2, multiple HLA loci, NOTCH4) and in established risk genes such as CACNA1C, TCF4, FURIN and MAD1L1, mirroring earlier reports [2].
Gene-Set Enrichment
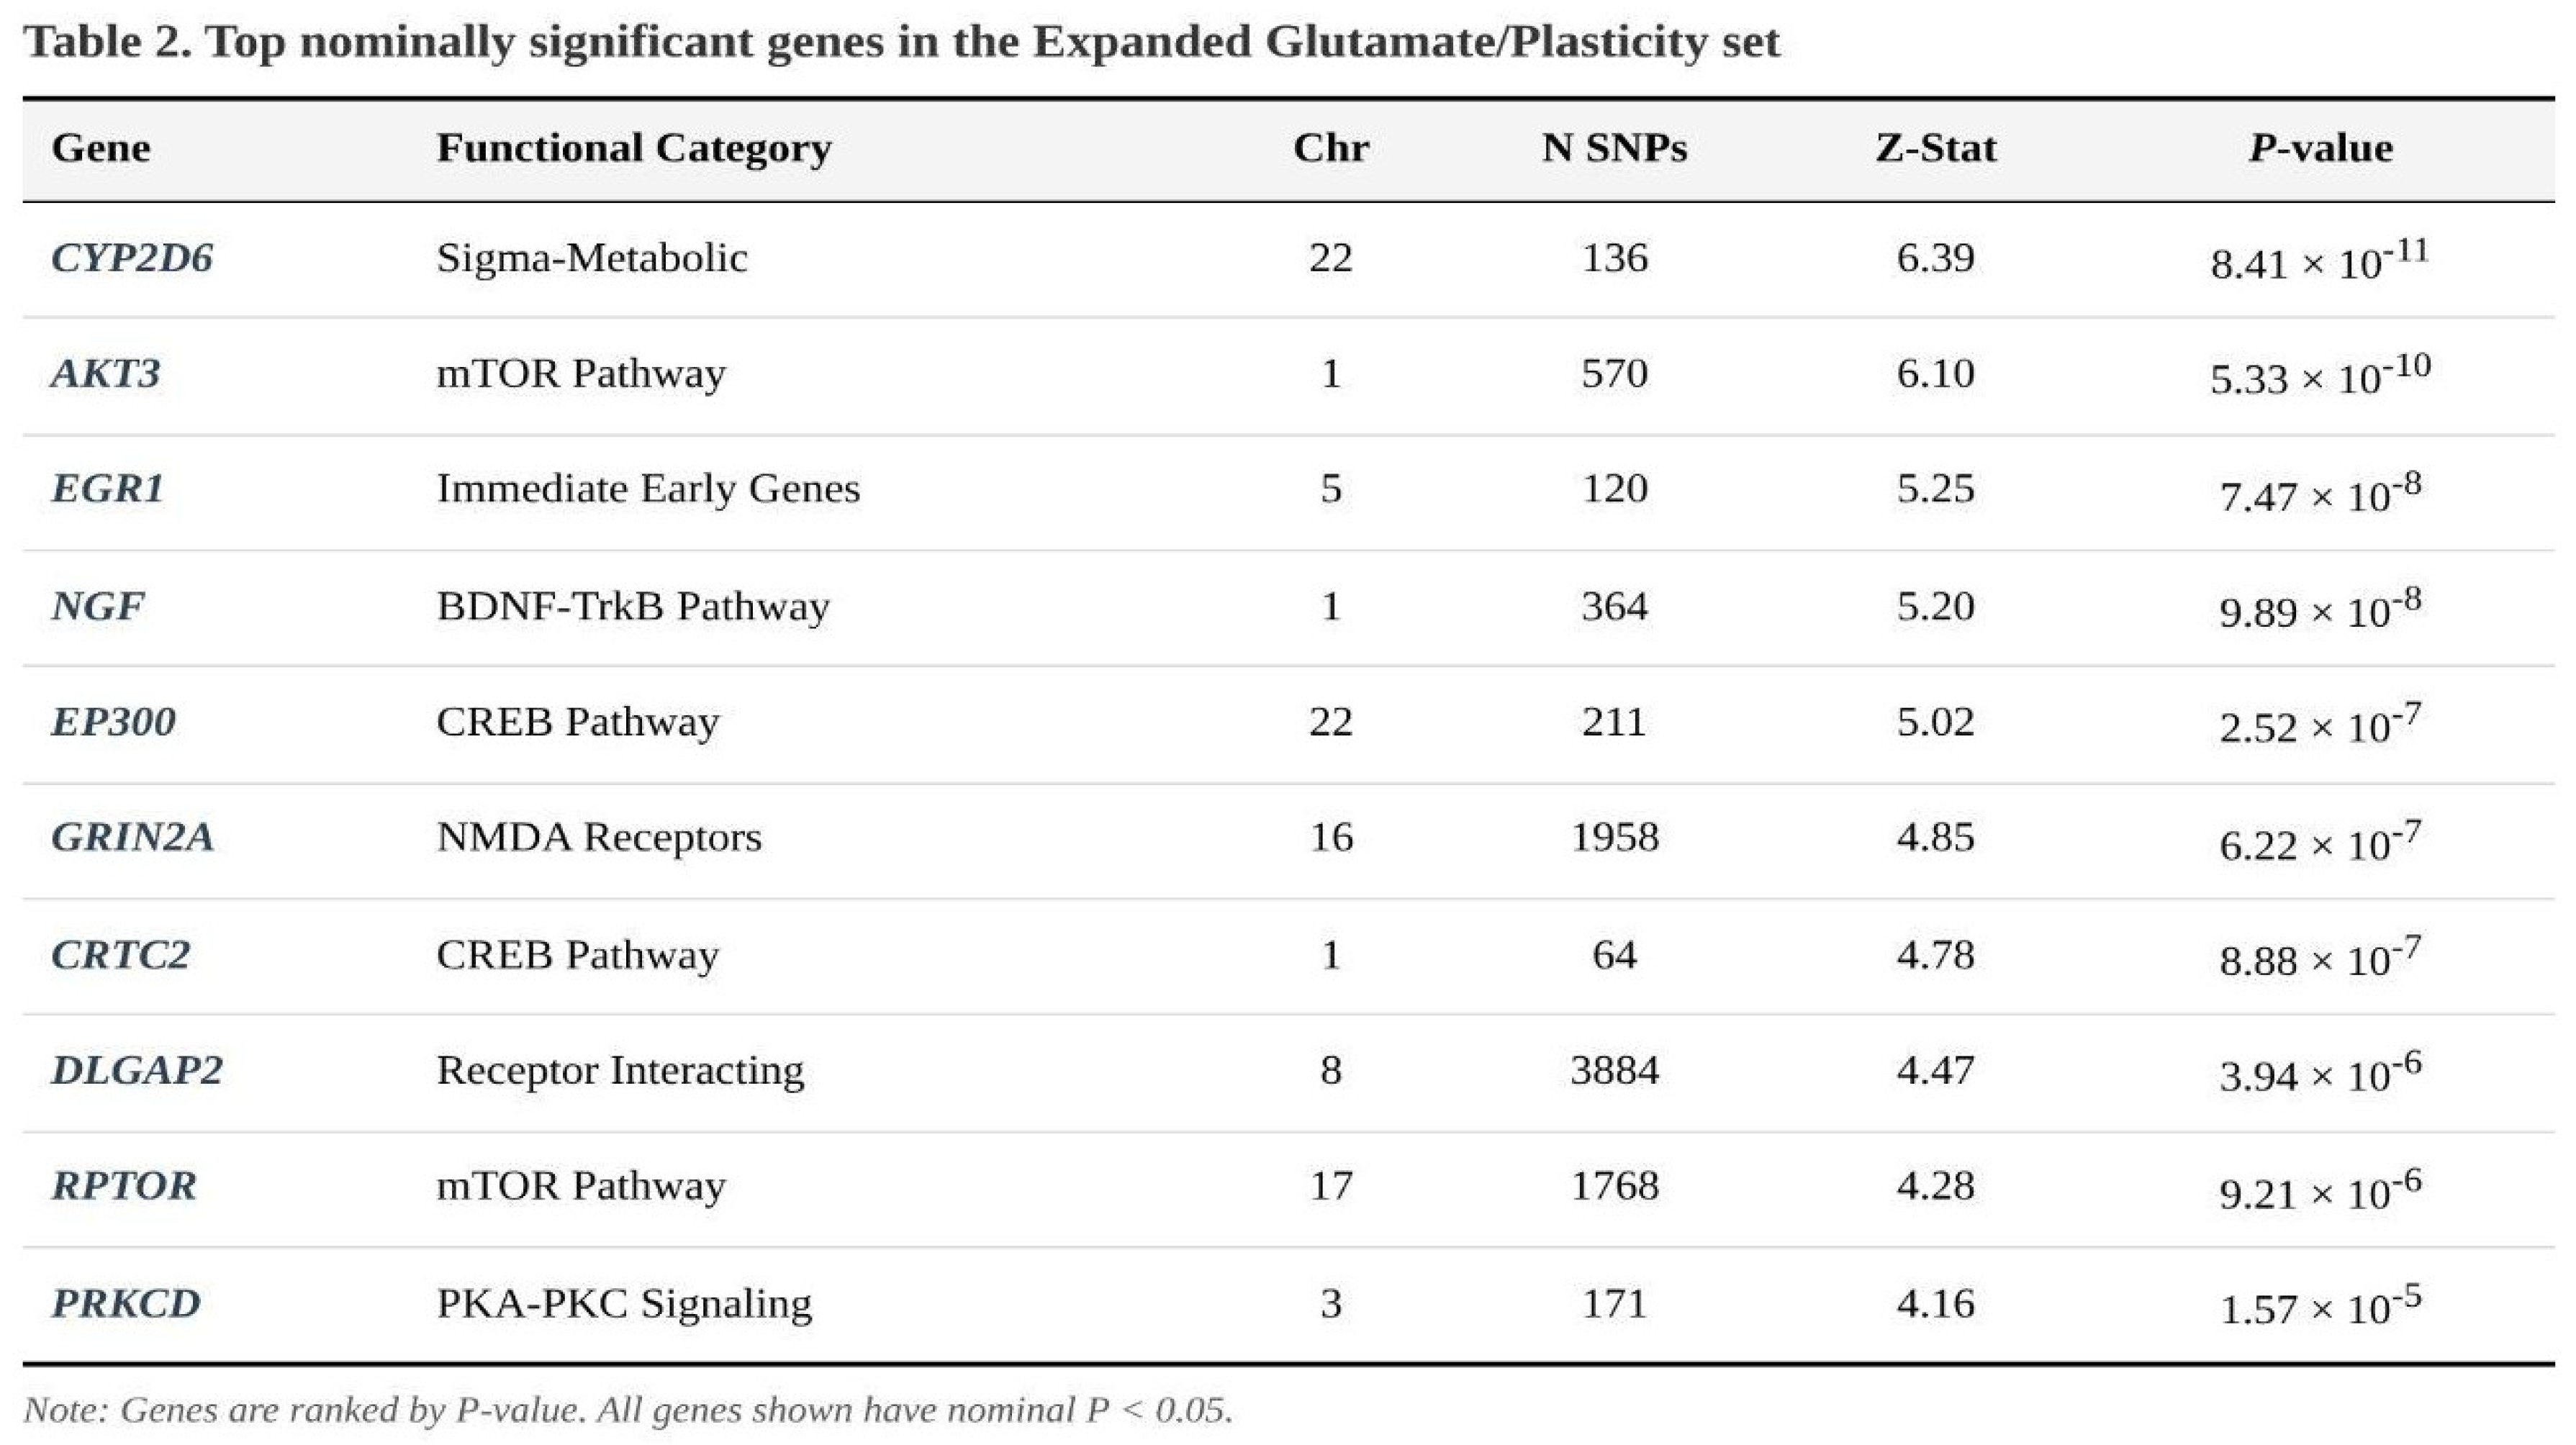
The 130-gene expanded glutamate/plasticity set was enriched for association (t = 2.97, p = 0.0033; Bonferroni p = 0.0134; FDR ≈ 0.013) (Table 1). Its mean Z-score (1.60) exceeded the genome-wide average (1.19). Sixty members of the expanded set reached nominal gene-level p < 0.05, and seven surpassed the genome-wide Bonferroni threshold. These included GRIN2A, AKT3 and several additional synaptic genes. CYP2D6 also met the gene-level threshold in our MAGMA run; however, given its location near complex LD regions and its primary role in drug metabolism, we treat this signal cautiously and note that it is not among the top-priority loci in the published PGC3 gene-level analyses [2]. Full gene-level statistics for all set members are provided in Appendix 1.
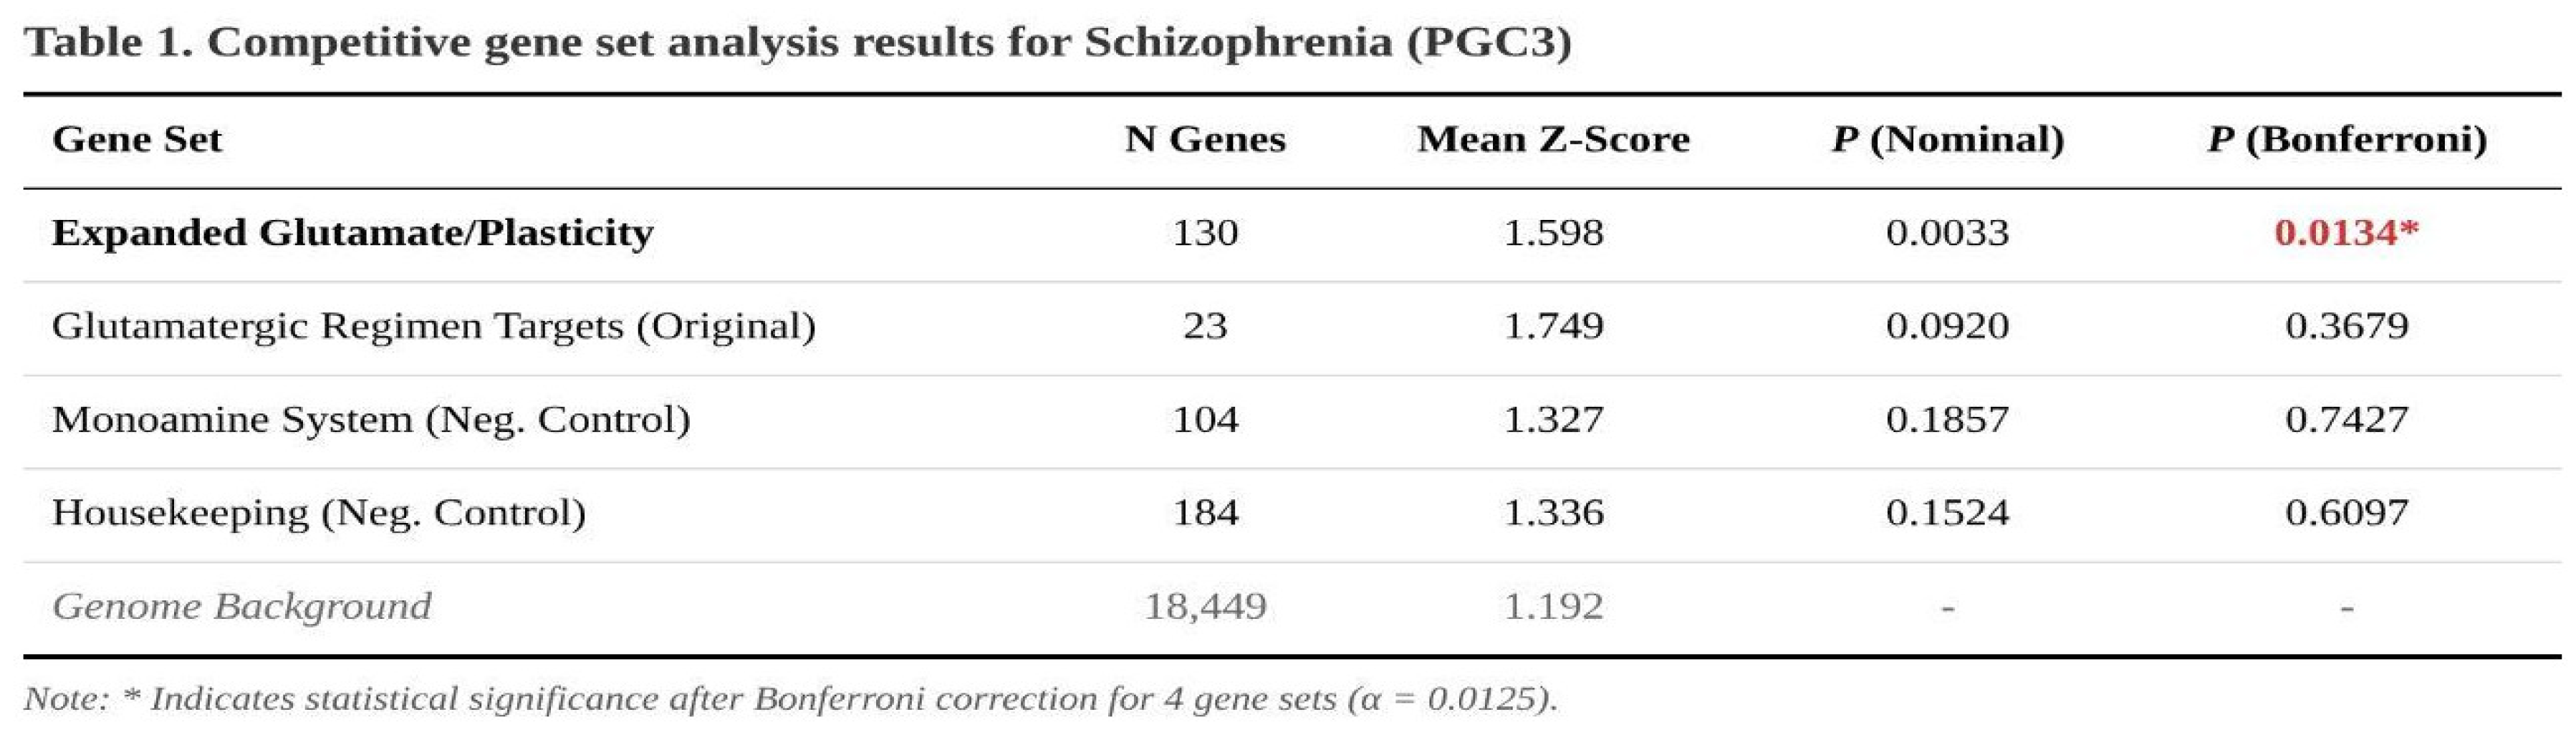
The 23-gene core set showed a non-significant trend (p = 0.092; Bonferroni p = 0.368). Eleven genes were nominally associated and two (CYP2D6, GRIN2A) met genome-wide criteria.
Neither control set displayed enrichment: monoamine (p = 0.186; Bonferroni p = 0.743) or house-keeping (p = 0.152; Bonferroni p = 0.610). A Kruskal–Wallis comparison of Z-score distributions across all four sets was not significant (H = 4.47, p = 0.215), but the glutamate-related sets nonetheless showed the highest central tendency.
Post-hoc inspection within the expanded set pointed to particularly strong contributions from sigma-metabolic genes, mTOR components, immediate-early genes, and BDNF–TrkB and CREB modules (Table 2).
Discussion
Gene-set analysis demonstrated significant enrichment of common risk variants within the expanded glutamatergic/plasticity pathway, even after stringent multiple-comparison correction. These results replicate and extend the PGC3 finding that common risk converges on synaptic biology and highlight broad glutamatergic signalling—beyond surface receptors—as a focal point of liability [2]. By contrast, neither the monoaminergic gene set nor housekeeping genes showed significant enrichment, reinforcing the specificity of the glutamatergic signal.
Notably, enrichment at the pathway level contrasted with patchy single-gene significance. For example, DRD2 showed an exceptionally strong individual association (p ≈ 10−14), yet the full monoamine pathway was not collectively enriched. This mirrors the longstanding paradox that, although D2 blockade is central to antipsychotic efficacy, GWAS implicate the broader dopamine pathway only sparsely [10]. Our results are consistent with this pattern: DRD2 itself shows a strong gene-level association, but the monoaminergic gene-set as a whole is not enriched. This suggests that common variant liability may be more diffusely distributed across glutamatergic/synaptic genes than across monoaminergic genes, even though individual monoaminergic genes such as DRD2 remain mechanistically crucial.
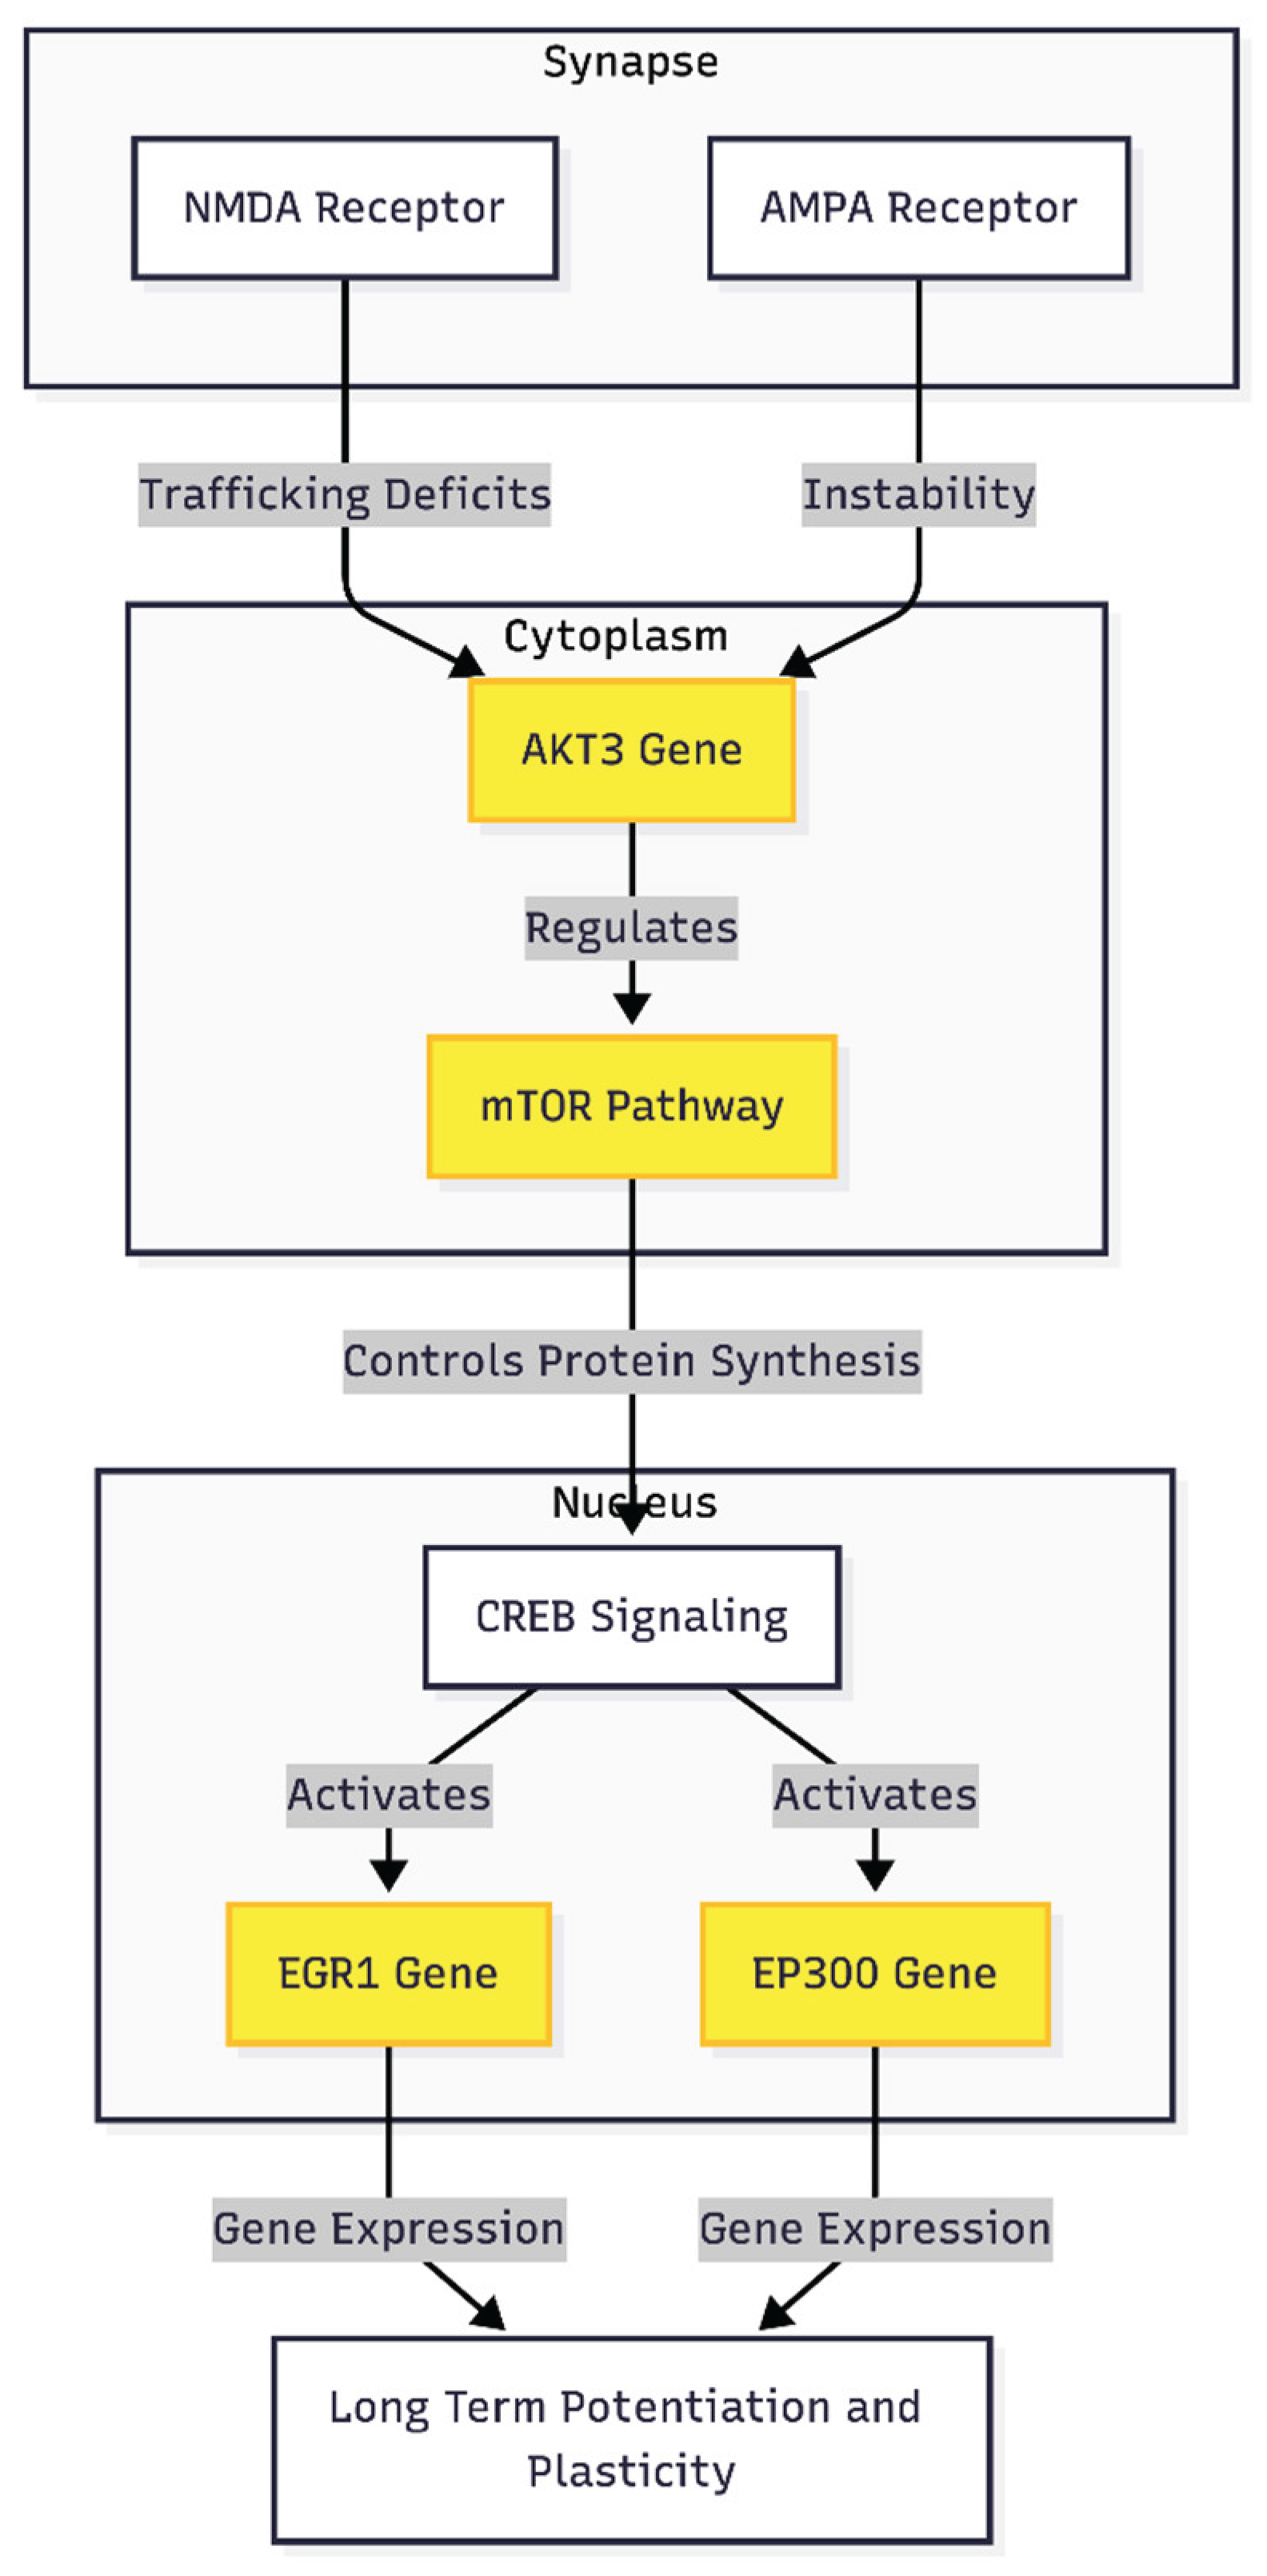
On the other hand, the larger glutamatergic set was important, while the smaller receptor-only set was not. Strong signals came from intracellular regulators like AKT3 (mTOR pathway), EGR1, EP300, and NGF, which are all important for plasticity that depends on protein synthesis [11]. When looked at with evidence from after death that NMDA was not working properly and AMPA trafficking was not working properly [4,5,12], the data suggest that genetic risk affects the molecular processes that are necessary for long-lasting changes in synapses (Figure 2).
Cognitive impairment, a major determinant of functional outcome, remains largely unaddressed by current dopamine-centric treatments [13]. Our findings argue for therapies aimed at enhancing synaptic plasticity. Compounds that modulate mTOR or CREB signalling, or that stabilise AMPA receptor insertion, could theoretically compensate for receptor-level deficits and improve cognition [14]. Furthermore, Pathway-specific polygenic scores derived from the expanded glutamate/plasticity set could, in principle, be used in future studies to stratify individuals in trials of glutamatergic or synaptic plasticity–targeting agents, but this remains to be empirically tested.
Strengths of this work include analysis of the largest schizophrenia GWAS to date, hypothesis-driven gene sets with relevant negative controls, and conservative statistics. Nonetheless, competitive MAGMA tests may under- or over-correct for genome-wide synaptic signals, and rare variant contributions were not examined. Future studies integrating common and rare variation and functional annotations may delineate precise molecular nodes for therapeutic targeting.
Funding
This research received no specific grant from any funding agency in the public, commercial, or not-for-profit sectors.
Ethics Declaration
Not applicable.
Conflicts of Interest
The regimen concerned (Cheung Glutamatergic Regimen, CGR) is not patented and is investigated for open-source usage as a low-cost flexible Ketamine-like psychiatric medication. Cheung Ngo Medical is a chain psychiatric clinic in Hong Kong owned by the corresponding author.
References
- Hilker, R; Helenius, D; Fagerlund, B; et al. Heritability of schizophrenia and schizophrenia spectrum based on the nationwide Danish Twin Register. Biological Psychiatry 2018, 83(6), 492–498. [Google Scholar] [CrossRef] [PubMed]
- Trubetskoy, V; Pardiñas, AF; Qi, T; et al. Mapping genomic loci implicates genes and synaptic biology in schizophrenia. Nature 2022, 604(7906), 502–508. [Google Scholar] [CrossRef] [PubMed]
- Krystal, JH; Karper, LP; Seibyl, JP; et al. Subanesthetic effects of the noncompetitive NMDA antagonist, ketamine, in humans: psychotomimetic, perceptual, cognitive, and neuroendocrine responses. Archives of general psychiatry 1994, 51(3), 199–214. [Google Scholar] [CrossRef] [PubMed]
- Clinton, SM; Meador-Woodruff, JH. Abnormalities of the NMDA receptor and associated intracellular molecules in the thalamus in schizophrenia and bipolar disorder. Neuropsychopharmacology 2004, 29(7), 1353–1362. [Google Scholar] [CrossRef] [PubMed]
- Hammond, JC; McCullumsmith, RE; Funk, AJ; et al. Evidence for abnormal forward trafficking of AMPA receptors in frontal cortex of elderly patients with schizophrenia. Neuropsychopharmacology 2010, 35(10), 2110–2119. [Google Scholar] [CrossRef] [PubMed]
- Bortolato, B; Miskowiak, KW; Köhler, CA; et al. Cognitive dysfunction in bipolar disorder and schizophrenia: A systematic review of meta-analyses. Neuropsychiatric Disease and Treatment 2015, 11, 3111–3125. [Google Scholar] [CrossRef] [PubMed]
- Li, W; Zhou, FC; Zhang, L; et al. Comparison of cognitive dysfunction between schizophrenia and bipolar disorder patients: A meta-analysis of comparative studies. Journal of affective disorders 2020, 274, 652–661. [Google Scholar] [CrossRef] [PubMed]
- Cheung, N. An oral “ketamine-like” NMDA/AMPA modulation stack restores cognitive capacity in a young man with schizoaffective disorder—Case report. Preprints 2025. [Google Scholar] [CrossRef]
- de Leeuw, CA; Mooij, JM; Heskes, T; et al. MAGMA: Generalized gene-set analysis of GWAS data. PLoS Computational Biology 2015, 11(4), e1004219. [Google Scholar] [CrossRef] [PubMed]
- Kendler, KS; Schaffner, KF. The dopamine hypothesis of schizophrenia: an historical and philosophical analysis. Philosophy, Psychiatry, & Psychology 2011, 18(1), 41–63. [Google Scholar] [CrossRef]
- Kelleher, RJ, 3rd; Govindarajan, A; Jung, HY; et al. Translational control by MAPK signaling in long-term synaptic plasticity and memory. Cell. 2004, 116(3), 467–479. [Google Scholar] [CrossRef] [PubMed]
- Dong, B; Yue, Y; Dong, H; et al. N-methyl-D-aspartate receptor hypofunction as a potential contributor to the progression and manifestation of many neurological disorders. Frontiers in molecular neuroscience 2023, 16, 1174738. [Google Scholar] [CrossRef] [PubMed]
- Vita, A; Gaebel, W; Mucci, A; et al. European Psychiatric Association guidance on treatment of cognitive impairment in schizophrenia. European Psychiatry 2022, 65(1), e57. [Google Scholar] [CrossRef] [PubMed]
- Hoeffer, CA; Klann, E. mTOR signaling: At the crossroads of plasticity, memory and disease. Trends in Neurosciences 2010, 33(2), 67–75. [Google Scholar] [CrossRef] [PubMed]
Figure 2.
Mechanistic Cascade. This figure maps the specific genes mentioned (AKT3, mTOR, EGR1, EP300) to their biological hierarchy, demonstrating how risk converges on intracellular regulation rather than just surface receptors.
Figure 2.
Mechanistic Cascade. This figure maps the specific genes mentioned (AKT3, mTOR, EGR1, EP300) to their biological hierarchy, demonstrating how risk converges on intracellular regulation rather than just surface receptors.
Disclaimer/Publisher’s Note: The statements, opinions and data contained in all publications are solely those of the individual author(s) and contributor(s) and not of MDPI and/or the editor(s). MDPI and/or the editor(s) disclaim responsibility for any injury to people or property resulting from any ideas, methods, instructions or products referred to in the content. |
© 2025 by the authors. Licensee MDPI, Basel, Switzerland. This article is an open access article distributed under the terms and conditions of the Creative Commons Attribution (CC BY) license (http://creativecommons.org/licenses/by/4.0/).
Copyright: This open access article is published under a Creative Commons CC BY 4.0 license, which permit the free download, distribution, and reuse, provided that the author and preprint are cited in any reuse.